

Cleaning Validation Change Control and Revalidation Triggers
Cleaning validation remains valid only while the assumptions, equipment, products, cleaning procedures, acceptance criteria, sampling methods, analytical methods, and operating conditions represented by the validation remain applicable. Change control provides the formal mechanism for determining whether a proposed or observed change affects that validated state and what additional evidence is needed before or after implementation.
A change does not automatically require cleaning revalidation. The appropriate question is whether the change can affect residue generation, cleaning difficulty, equipment cleanability, carryover risk, cleaning-process effectiveness, sampling representativeness, analytical capability, or another assumption supporting the original validation. Where those elements remain adequately covered by existing evidence, documented assessment may be sufficient. Where they change materially, targeted verification or broader revalidation may be necessary.
This distinction prevents two opposite errors: repeating full cleaning validation for every minor change regardless of impact, or approving significant changes administratively without determining whether the original validation still represents the revised process.
FDA’s cleaning-validation inspection guide expects the manufacturer’s validation program to define when revalidation will be required. ICH Q7 provides the broader lifecycle principle that validated systems and processes should be periodically evaluated and that, where no significant changes have occurred and quality review demonstrates consistent acceptable performance, revalidation is normally unnecessary.
Change Control Protects the Validated State
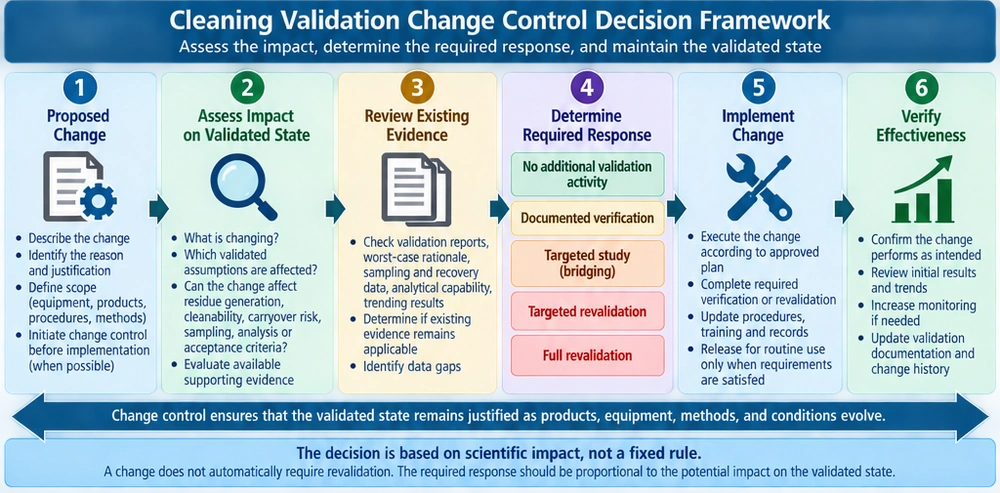
Cleaning-validation change control should begin before implementation whenever the change is planned. The assessment should identify what is changing, determine which validated assumptions are affected, evaluate the available supporting evidence, establish any required verification or revalidation, and define the conditions for release of the revised process.
The logic is:
Proposed change → cleaning-validation impact assessment → existing evidence review → determine additional evidence → implement under change control → verify effectiveness → update validation documentation
The objective is not simply to assign a risk score. The assessment should reach a technically defensible conclusion about whether the cleaning process remains represented by the approved validation evidence.
Cleaning Validation Program Strategy, Scope, and Lifecycle should establish the overall validation framework, while this article defines how changes are evaluated after that framework is established.
Regulatory and Lifecycle Basis
21 CFR 211.67 — Equipment Cleaning and Maintenance requires written procedures describing equipment cleaning methods, materials, disassembly and reassembly where necessary, cleaning schedules, and inspection for cleanliness. Changes affecting those elements therefore can affect the basis on which the equipment cleaning process was validated.
21 CFR 211.100 — Written Procedures; Deviations requires changes to written production and process-control procedures to be drafted, reviewed, and approved by the appropriate organizational units and the quality control unit. It also requires deviations from written procedures to be recorded and justified.
For automated cleaning systems, 21 CFR 211.68 — Automatic, Mechanical, and Electronic Equipment requires appropriate control of computerized systems and routine checks designed to assure proper equipment performance. Changes to automated cleaning recipes, software, sequencing, or parameter control can therefore have both computerized-system and cleaning-validation implications.
ICH Q7 states that validated systems and processes should be periodically evaluated and that revalidation is normally unnecessary where no significant changes have occurred and review confirms continued acceptable operation. Its cleaning-validation section further states that validation should reflect actual equipment-use patterns and should define equipment, cleaning procedures, cleaning materials, acceptance levels, controlled parameters, analytical methods, and sampling. These are therefore logical categories for change-impact assessment.
Not Every Change Is a Revalidation Trigger
A change should first be evaluated for validation impact rather than automatically labeled a revalidation trigger.
For example, replacing a non-product-contact equipment nameplate with an equivalent part generally has no plausible effect on cleaning effectiveness. Replacing a product-contact gasket with a different material can affect residue adhesion, cleanability, chemical compatibility, sampling recovery, and surface representation and therefore requires a more substantive assessment.
Similarly, correcting a typographical error in a cleaning instruction is different from changing detergent concentration or reducing wash time.
The level of validation response should follow the technical impact of the change.
A Practical Revalidation Decision Structure
The change-control assessment should determine whether the change:
- affects a previously validated worst-case condition;
- introduces a new or more severe cleaning challenge;
- alters an approved cleaning parameter or procedure;
- changes product-contact equipment or materials;
- changes the acceptance limit or its calculation;
- changes sampling representativeness or recovery;
- changes analytical capability;
- modifies automation or recipe logic affecting cleaning;
- invalidates product or equipment grouping assumptions;
- is associated with evidence of deteriorating cleaning performance.
If none of these conditions is affected and the rationale is technically supported, existing validation may remain applicable. If one or more are affected, additional evidence should be defined according to the nature and magnitude of the change.
New Product Introduction
A new product introduced into shared equipment is a significant cleaning-validation assessment point because it can create a challenge not represented by the existing product matrix.
The assessment should consider the product’s HBEL or PDE, formulation, potency, solubility, cleanability, residue characteristics, dosage, manufacturing batch size, processing conditions, campaign length, dirty hold time, and interaction with product-contact surfaces. The potential sequence with subsequent products should also be evaluated because the applicable MACO can depend on both the previous and next product.
The new product should then be compared with the products already represented in Worst-Case Product, Equipment, and Cleaning Condition Selection.
If the existing worst-case product remains more challenging for both carryover risk and cleanability, the new product may be incorporated through documented bracketing or grouping without repeating full cleaning validation. If the new product creates a more restrictive HBEL-derived limit, is more difficult to remove, or introduces a different residue mechanism, targeted or broader validation may be required.
A current FDA warning letter from August 2026 specifically requested that a manufacturer describe the change-management steps required before introducing new manufacturing equipment or a new product and called for improvements to cleaning validation incorporating worst-case toxicity, potency, solubility, equipment cleanability, difficult swab locations, and maximum hold times.
Changes to HBEL, PDE, or ADE
A revised health-based exposure limit can materially change the residue acceptance criteria without any physical change to the cleaning process.
Health-Based Exposure Limits for Cleaning Validation should identify the toxicological source and lifecycle of the HBEL. When that value changes, the impact assessment should propagate the new value through Maximum Allowable Carryover (MACO) in Cleaning Validation and Cleaning Validation Acceptance Criteria and Residue Limits.
The assessment should determine whether the revised limit affects the MACO, surface criterion, swab or rinse acceptance limit, analytical LOQ, previous worst-case rankings, or interpretation of historical validation results.
A lower HBEL does not automatically require repeating cleaning validation. If historical results remain comfortably below the recalculated acceptance criterion and the analytical procedure remains capable, existing validation evidence may continue to support the process. If the new limit is below demonstrated cleaning performance or below reliable analytical capability, additional work becomes necessary.
An increased HBEL likewise does not require relaxation of an existing tighter cleaning criterion. The site may retain a more restrictive operational limit when it remains appropriate and achievable.
Product Formulation Changes
A formulation change can affect cleanability even when the API and toxicological limit remain unchanged. Changes in binders, polymers, oils, coatings, excipients, solids concentration, pH, viscosity, or processing conditions can change residue adhesion, solubility, drying behavior, or interaction with equipment surfaces.
A reformulated product should therefore be compared with the validated soil or product represented in Cleaning Procedure Development and Efficacy Studies.
The key question is whether the revised formulation creates a cleaning challenge equal to or less than the validated condition. Where development or laboratory evidence demonstrates equivalence or lesser challenge, targeted documentation may be sufficient. Where the formulation creates a materially different soil or harder-to-remove residue, additional efficacy work and cleaning verification may be required before concluding that the existing validation remains applicable.
Changes to Product Strength or Dose
A product strength change can affect the cleaning-validation program in more than one way. It may change residue loading during manufacture, formulation composition, maximum daily dose, or the product’s role as a subsequent product in MACO calculations.
A strength increase does not automatically mean the product is harder to clean, but the change should be evaluated for its effect on the applicable carryover calculations and the physical residue challenge.
Where multiple strengths were already bracketed during validation, the assessment should determine whether the new strength remains within the validated range and formulation envelope.
Batch-Size Changes
Batch-size changes can affect both cleaning challenge and carryover calculations.
For a PDE-based MACO calculation, the minimum applicable subsequent-product batch size is normally the more restrictive batch-size condition. Introduction of a smaller subsequent batch may therefore lower the allowable carryover.
A larger previous-product batch or altered process fill can also increase residue loading on equipment even if the calculated carryover limit is unchanged.
The assessment should consider both sides of the problem: allowable carryover and amount or distribution of soil requiring removal.
Campaign-Length Changes
Extending the permitted campaign can increase residue accumulation and can change the condition of equipment at the time cleaning begins. Residue may build progressively at gaskets, filters, transfer lines, outlets, or other retention points.
If campaign length was part of the validated worst-case condition, increases beyond that range should trigger assessment of whether additional cleaning-development work, verification, or revalidation is necessary.
A reduced campaign usually represents a lower cleaning challenge, but the impact should still be documented if campaign assumptions are part of approved cleaning grouping or validation documentation.
Dirty Hold Time Changes
Dirty hold time can materially affect residue drying, adhesion, crystallization, microbial growth, and cleaning difficulty. Increasing the maximum permitted dirty hold beyond the validated condition is therefore a significant cleaning-validation change.
Dirty Hold Time and Clean Hold Time Studies should define the established dirty-hold claim and supporting evidence.
An extension should normally be supported before implementation through appropriate studies demonstrating that the cleaning process remains effective at the longer interval. If the new condition becomes more challenging than that represented during formal cleaning validation, targeted revalidation may be needed.
A shorter dirty hold generally does not create a greater residue-removal challenge, although operational or microbiological consequences should still be considered.
Clean Hold Time Changes
An extended clean hold does not normally challenge removal of the previous residue because cleaning has already occurred, but it can affect the continued suitability of cleaned equipment through microbial proliferation, environmental contamination, moisture retention, or storage conditions.
The assessment should therefore focus on whether the clean equipment remains acceptable throughout the revised storage interval and whether inspection, sanitization, or re-cleaning requirements need to change.
Where the clean-hold claim is materially extended, the supporting hold-time study may need to be expanded rather than repeating the chemical-residue cleaning validation itself.
Equipment Modification
Product-contact equipment changes require evaluation of both cleanability and representativeness of existing validation evidence.
Examples include changes to vessels, pumps, valves, agitators, spray devices, transfer piping, filter housings, filling pathways, hoses, gaskets, dead-leg configuration, drain points, surface finish, and product-contact materials.
The assessment should determine whether the modification changes:
- product-contact surface area;
- residue-retention geometry;
- cleaning solution coverage;
- flow or turbulence;
- drainage;
- manual accessibility;
- disassembly;
- spray shadowing;
- sampling accessibility;
- surface material;
- recovery characteristics.
FDA’s cleaning-validation guide specifically emphasizes equipment design, piping, valves, cleaning access, and process details because these elements can materially affect cleaning effectiveness.
Like-for-Like Replacement
A documented like-for-like replacement does not necessarily require revalidation. The assessment should confirm that the replacement is genuinely equivalent in the characteristics relevant to cleaning.
For a product-contact component, equivalence may include material, geometry, surface finish, dimensions, functionality, orientation, and cleanability.
Calling a replacement “like-for-like” does not make it so. A gasket of the same dimensions but different elastomer composition can have different chemical resistance, surface interaction, and sampling recovery.
Where the replacement is demonstrated to be equivalent and the cleaning mechanism is unchanged, existing validation can normally remain applicable.
Changes in Product-Contact Materials
A new surface material can affect residue adhesion, corrosion resistance, cleaning-chemical compatibility, and sampling recovery.
If stainless steel is replaced with a polymer, elastomer, coating, glass surface, or different alloy, the impact assessment should consider both cleaning effectiveness and the applicability of existing Swab and Rinse Recovery Studies for Cleaning Validation.
The change may require a new recovery study even when full cleaning revalidation is unnecessary.
This illustrates why change control should determine the specific evidence affected rather than default to either “no action” or “repeat everything.”
Cleaning Chemistry Changes
Changes to detergent, solvent, concentration, formulation, supplier, or cleaning-agent composition can directly affect residue removal and should receive formal assessment.
The cleaning chemistry selected during Cleaning Procedure Development and Efficacy Studies is part of the validated cleaning mechanism. Replacing one detergent with another should not be justified solely because the new material has a similar commercial description or pH.
The assessment should consider soil compatibility, concentration range, temperature dependence, surface compatibility, rinsability, cleaning-agent residue, and analytical detection where applicable.
A substantially different cleaning chemistry may require development or comparative efficacy studies followed by targeted cleaning validation.
Cleaning-Parameter Changes
Changes to cleaning parameters can alter the validated cleaning mechanism even when equipment and detergent remain unchanged.
Relevant parameters can include wash time, temperature, detergent concentration, flow, pressure, mechanical action, spray conditions, number of rinses, rinse volume, conductivity endpoint, pH endpoint, manual scrubbing duration, soaking time, and drying conditions.
A proposed parameter change should be evaluated against the range or worst-case condition represented during validation. If the new value remains inside the validated range, no new validation may be required. If it extends beyond the validated range in a direction that could reduce cleaning effectiveness, additional evidence is needed.
A shorter wash time, lower temperature, lower detergent concentration, or reduced mechanical action generally warrants more attention than a change that increases cleaning severity, but increased severity can introduce other concerns such as equipment damage, residue generation, detergent carryover, or material incompatibility.
Cleaning Revalidation Trigger Impact Matrix
Cleaning-validation changes should be assessed according to how they affect the assumptions and evidence supporting the validated state. The presence of a change does not automatically require revalidation; the appropriate response depends on whether the change affects patient-safety limits, cleaning difficulty, equipment or process performance, sampling representativeness, analytical capability, or the applicability of existing validation evidence.
| Change | Examples | Potential Cleaning-Validation Impact | Typical Validation Response |
|---|---|---|---|
| New product or formulation | New API, dosage form, excipients, coating, viscosity, product strength | May introduce a lower HBEL, more difficult residue, different soil characteristics, or a new worst-case condition | Assess against existing worst-case grouping; targeted study or revalidation if not adequately represented |
| HBEL / PDE / ADE change | Revised toxicological assessment, new dose or patient population | Changes MACO, surface/sample limits, analytical requirements, and potentially worst-case ranking | Recalculate limits and assess historical data; targeted verification if existing evidence does not support the revised limit |
| Equipment or product-contact surface change | New vessel, valve, gasket, hose, pump, surface material or finish | May affect cleanability, drainage, residue retention, surface area, sampling access, or recovery | Document equivalence for true like-for-like changes; recovery study, targeted verification, or revalidation where cleaning characteristics change |
| Cleaning chemistry change | New detergent or solvent, concentration change, supplier/formulation change | May alter residue removal, rinsability, equipment compatibility, or cleaning-agent residue | Comparative efficacy or bridging study; targeted revalidation where the cleaning mechanism changes materially |
| Cleaning parameter change | Time, temperature, flow, pressure, detergent concentration, number of rinses, manual contact time | Can alter cleaning effectiveness or move operation outside the validated range | No additional validation if clearly within validated range; targeted verification or revalidation when the validated boundary is extended |
| Software or cleaning-recipe change | CIP sequence, valve logic, setpoints, timers, dosing, alarms, interlocks | Can change how the physical cleaning process is executed | Assess both computerized-system and cleaning impact; targeted verification or revalidation when cleaning execution changes |
| Sampling or recovery change | New swab, rinse solvent, sample location, swab area, extraction volume, recovery factor | May affect representativeness, recovery, sample concentration, or comparability with historical data | Usually targeted recovery or bridging study; reassess acceptance calculations and sampling rationale |
| Analytical-method change | New HPLC/UPLC/TOC method, revised LOQ, range, preparation, reporting convention | May affect quantitative capability and comparability with previous validation results | Analytical bridging or method validation; cleaning revalidation only when existing evidence can no longer support the required criterion |
| Disinfectant or sanitizer change | New agent, concentration, contact time, addition/removal from product-contact cleaning cycle | May affect residues, rinsing, surface compatibility, or microbiological control | Assess whether change affects product-contact cleaning; perform efficacy or residue studies as applicable |
| Dirty hold or campaign change | Longer dirty hold, increased campaign length, increased residue loading | May increase drying, adhesion, accumulation, or cleaning difficulty | Hold-time or cleaning-efficacy study; targeted revalidation if the new condition exceeds the validated worst case |
| Maintenance or repair | Welding, polishing, valve replacement, gasket replacement, spray-device repair | May change surface condition, geometry, coverage, drainage, or sampling recovery | Return-to-service assessment for equivalent repair; targeted verification when product-contact or cleaning characteristics change |
| Adverse performance trend or failure | Increasing residue results, repeated recleaning, visual failures, recurring deviations | May indicate loss of process capability despite no planned change | Investigation, increased verification, CAPA, and targeted or broader revalidation depending on cause and extent |
The matrix should be used as a decision aid rather than as a fixed revalidation rule. The final response should be based on the specific validated assumptions affected, the strength of existing supporting evidence, and the magnitude of the change, with the outcome ranging from documented no-impact justification through targeted verification or bridging to targeted or full revalidation.
Manual Cleaning Procedure Changes
Manual cleaning procedures depend heavily on sequence, access, tools, mechanical action, solution preparation, contact time, rinsing, and operator technique. Changes to these instructions can therefore have direct validation impact.
Examples include changing brush type, eliminating equipment disassembly, changing the number of passes, revising soaking time, modifying detergent preparation, changing rinse technique, or altering the order of cleaning steps.
The assessment should determine whether the revised procedure produces an equivalent or more robust cleaning mechanism. Where operator technique is significant, additional training and targeted verification may be necessary even when the change does not justify full revalidation.
Automated Cleaning Recipe and Software Changes
Automated CIP and washer systems can control recipe steps, valves, pumps, detergent addition, temperatures, durations, rinse endpoints, alarms, and sequence logic. A software change can therefore become a cleaning-validation change when it alters how the physical cleaning process operates.
The assessment should distinguish between changes that affect only system administration or presentation and changes that can alter cleaning execution.
A display-format correction may have no cleaning-validation impact. Changes to recipe logic, timer calculation, valve sequencing, conductivity endpoint, pump control, detergent dosing, interlocks, or parameter ranges can directly affect the validated cleaning process.
21 CFR 211.68 requires appropriate controls over automated and computerized equipment and records and written programs intended to assure proper equipment performance. Cleaning-related software changes should therefore be assessed through both the computerized-system change process and the cleaning-validation impact process where applicable.
Recipe Setpoint Versus Validated Range
A change to a routine setpoint does not necessarily require revalidation when the new setpoint remains within a previously validated operating range.
For example, if validation demonstrated acceptable cleaning at a minimum wash temperature of 65°C and routine operation was set at 70°C, changing the routine setpoint to 68°C may remain covered provided the validated minimum and other relevant conditions remain unchanged.
By contrast, changing the minimum allowable temperature to 60°C extends the cleaning process beyond the established validated boundary and requires evidence supporting that condition.
The assessment should therefore compare the proposed operating range with the validated range rather than focusing only on the current production setpoint.
Sampling-Location Changes
Sampling locations are part of the cleaning-validation evidence. Removing or replacing a location should therefore be justified by the cleaning-risk assessment rather than convenience.
If an equipment modification makes the original sampling location inaccessible, Cleaning Validation Sampling Strategy and Worst-Case Locations should be reassessed to determine how the same cleaning-risk mechanism will be represented.
Introducing a new sampling location does not automatically require revalidation, but recovery applicability, surface material, area, acceptance calculations, and analytical method should be evaluated.
A worst-case location should not be removed simply because historical results have consistently passed. Those results may demonstrate that the cleaning process is effective precisely because the location remains controlled.
Swab or Rinse Method Changes
Changing swab material, sampling solvent, swab area, rinse solvent, rinse volume, extraction volume, sampling pattern, contact time, or collection configuration can affect the measured result even when equipment cleanliness is unchanged.
Swab Sampling for Cleaning Validation and Rinse Sampling for Cleaning Validation should define the validated sampling methods.
The impact assessment should determine whether existing recovery data remain applicable and whether the analytical criterion needs recalculation.
A change from 10 mL to 20 mL swab extraction volume, for example, halves the theoretical analytical concentration associated with the same surface residue limit. This may affect method sensitivity without altering the actual cleaning requirement.
Recovery-Method Changes
Recovery factors are part of the measurement system. A new swab, solvent, surface material, sampling pattern, or rinse procedure can invalidate the existing recovery factor.
Swab and Rinse Recovery Studies for Cleaning Validation should therefore be included in change impact whenever the sampling interface changes.
In many cases the appropriate response is a targeted recovery or bridging study rather than repeated cleaning validation.
If the revised recovery substantially changes historical result interpretation or causes the method no longer to support the established acceptance criterion, broader impact assessment is required.
Analytical Method Changes
Changes to chromatography, TOC, conductivity, sample preparation, analytical range, LOQ, calculation, instrumentation, or reporting convention can affect the ability to demonstrate compliance with the cleaning criterion.
A new analytical method can be superior without requiring the cleaning process itself to be revalidated. The site should determine whether the new method has adequate comparability with historical evidence and whether results before and after the change can be interpreted consistently.
If a substantially more sensitive method begins detecting previously unquantifiable residue, that finding should be evaluated scientifically. FDA does not expect cleaning limits to become progressively lower simply because analytical sensitivity improves; the established cleaning limit should remain based on a justified safety and quality rationale. The method must be capable of verifying that criterion.
Analytical LOQ Changes
A changed LOQ should be compared with the sample concentration corresponding to the approved cleaning acceptance criterion.
If the revised method remains quantitatively capable at the required level, no cleaning revalidation may be necessary. If the LOQ becomes too high to support the established criterion, the analytical method is no longer suitable for the intended cleaning-validation decision.
The correct response is to restore analytical capability or modify the sampling method—not to weaken the patient-safety-based acceptance limit.
Acceptance-Criterion Changes
Changes to MACO, surface limits, swab criteria, rinse criteria, detergent limits, or visual requirements should trigger assessment of historical validation evidence.
A tighter criterion may still be supported by existing results. If all prior validation data remain below the new limit and the analytical procedure was capable at that level, repeating validation may add little scientific value.
If historical results cannot demonstrate compliance with the revised criterion, additional verification or revalidation may be necessary.
A less restrictive criterion should likewise be justified through the approved toxicological or quality-risk basis rather than simply adopted because it simplifies routine operation.
Disinfectant or Sanitizer Changes
Disinfectant and sanitizer changes require careful boundary definition because not every disinfection change is a cleaning-validation change.
Where a sanitizer or disinfectant is an integral part of the product-contact equipment cleaning cycle, changing the agent, concentration, contact time, sequence, or application method can affect the validated equipment condition and should be assessed through cleaning change control.
Where the change concerns cleanroom or controlled-area surface disinfection rather than product-contact cleaning, the primary validation impact may fall under Disinfectant Efficacy Studies for GMP Cleanrooms and Controlled Areas rather than chemical-residue cleaning validation.
The assessment should consider antimicrobial efficacy, residues, compatibility with surfaces, required rinsing, interaction with cleaning agents, and any impact on subsequent manufacturing.
This distinction prevents cleanroom disinfectant qualification and product-contact cleaning validation from being treated as the same program.
Maintenance and Repair
Maintenance can alter surfaces or equipment configuration even when no formal engineering project is initiated.
Relevant activities include welding, polishing, gasket replacement, pump repair, valve replacement, spray-device maintenance, hose replacement, surface repair, sensor replacement, and piping modification.
After maintenance, the assessment should determine whether product-contact surfaces, cleaning coverage, drainage, configuration, sampling accessibility, or recovery have changed.
Routine preventive maintenance using equivalent parts may require only documented return-to-service checks. Major repair affecting product-contact geometry or surface condition may require targeted cleaning verification or revalidation.
Utilities Affecting Cleaning
Changes to utilities used during cleaning can also affect cleaning performance. Examples include changes in purified-water quality, WFI supply, clean steam where applicable, water temperature, pressure, flow capacity, compressed gas used for drying, or detergent supply systems.
The cleaning-validation impact should be based on whether the utility characteristic is important to the validated cleaning mechanism.
A utility modification that maintains all validated conditions may have no cleaning revalidation impact. A change that reduces available flow below the validated minimum or alters rinse-water quality may directly affect cleaning effectiveness.
Changes Caused by Deviations or Performance Trends
Not all triggers arise from planned changes. Ongoing Cleaning Verification and Performance Trending may identify increasing residue levels, repeated visual findings, rising variability, recleaning, or recurring cleaning deviations.
Such evidence should trigger reassessment of whether the cleaning process still operates within the validated state.
Similarly, Cleaning Validation Deviations, Failures, and Investigations may identify a root cause that requires procedure, equipment, parameter, or analytical changes.
The resulting CAPA should pass through change control, and the investigation should define what verification or revalidation is necessary to demonstrate that the corrected process is effective.
Change Impact Should Consider Historical Data
A strong impact assessment does not rely only on theoretical risk. Existing cleaning data can provide important evidence about process capability and margin.
For example, if an HBEL change reduces a surface criterion from 2.0 to 1.5 µg/cm² while five years of worst-case results remain below 0.2 µg/cm², existing evidence may strongly support continued validation without repeating the original study.
Conversely, if routine results frequently approach the original limit, even a modest tightening can materially affect the validated state.
Historical performance should therefore be interpreted together with the technical nature of the change.
Determine the Required Validation Response
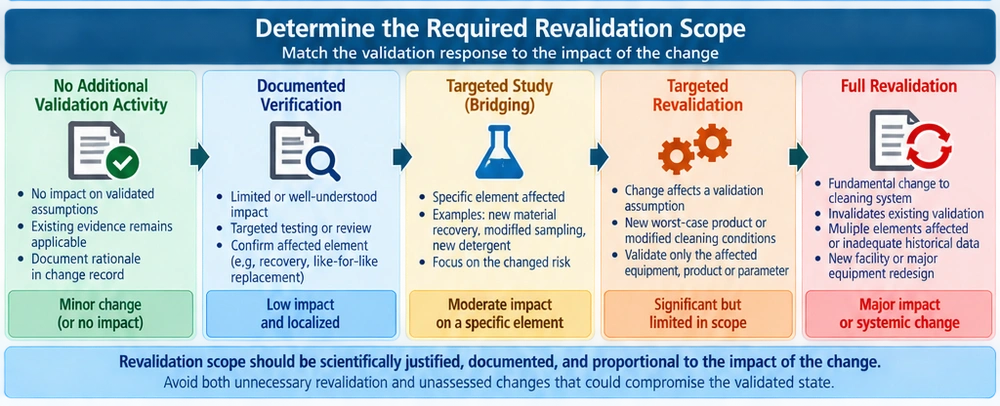
The outcome of the change assessment should be explicit. A practical hierarchy is:
- No additional validation activity — existing evidence remains fully applicable.
- Documented verification — confirm an aspect of the change without repeating formal validation.
- Targeted study or bridging — evaluate a specific affected element such as recovery, new surface material, new product cleanability, or modified parameter.
- Targeted cleaning revalidation — execute cleaning validation against the affected equipment, product, condition, or location.
- Broader/full revalidation — repeat a more comprehensive validation where the change materially alters the cleaning process or invalidates the existing validation basis.
The decision should reflect the actual affected evidence rather than use a predetermined rule based solely on change classification.
Targeted Verification
Targeted verification is appropriate when the original validation remains substantially applicable but one specific element requires confirmation.
Examples include testing recovery on a new gasket material, verifying cleaning after a like-for-like but product-contact repair, assessing a new low-risk product already covered by a validated group, or confirming performance after a limited parameter adjustment that remains close to the validated range.
The protocol or study should clearly identify what uncertainty the targeted work is intended to resolve.
Once the required evidence is obtained, the change record should state why broader revalidation is unnecessary.
Targeted Revalidation
Targeted revalidation goes further than a bridging study and directly challenges the modified cleaning process under the affected condition.
Examples can include a new worst-case formulation, modified valve geometry, substantially changed detergent concentration, revised manual cleaning step, or extension of dirty hold beyond the previously validated claim.
The study should focus on the changed risk rather than mechanically repeat all prior validation activities that remain unaffected.
Full Revalidation
Full or broad revalidation becomes appropriate when the change fundamentally alters the cleaning system or when multiple original validation assumptions are no longer applicable.
Examples may include major equipment redesign, replacement of the cleaning chemistry and cycle, substantial automated recipe redevelopment, major product portfolio changes that invalidate existing worst-case grouping, or repeated failures demonstrating that the prior cleaning process is no longer reproducible.
Full revalidation can also become necessary when poor historical documentation prevents the site from determining whether existing validation adequately represents the revised process.
Define Requirements Before Implementing the Change
Where additional validation work is required, the change record should identify it before implementation whenever practical.
The change plan should define the required protocol, sampling, analytical method, acceptance criteria, affected equipment or products, responsible functions, prerequisite activities, and release condition.
This prevents the change from entering routine use before the organization has established whether the revised cleaning process is adequately controlled.
Temporary or emergency changes should still receive documented risk assessment and retrospective completion of required validation activities according to the site’s quality system.
Effectiveness Verification
A change can be technically justified yet fail during routine implementation. Effectiveness verification should therefore confirm that the implemented change performs as expected.
Evidence can include targeted residue testing, visual inspection, process-parameter review, recovery data, initial-cycle monitoring, trend review, or other measurements appropriate to the change.
For higher-risk changes, the ongoing verification frequency may be temporarily increased after implementation until sufficient evidence demonstrates stable performance.
This links formal change control directly with Ongoing Cleaning Verification and Performance Trending.
Update the Validation Documentation
Approved changes should be reflected in the controlled documentation supporting cleaning validation.
Depending on the change, updates may be required to the cleaning SOP, validation matrix, equipment surface-area calculations, HBEL/MACO database, acceptance criteria, worst-case rationale, sampling map, analytical method, recovery study, cleaning recipe, validation report, and periodic-review records.
The validation system should not depend on an old protocol that describes equipment, parameters, or products no longer used.
Traceability should allow a reviewer to understand the current validated configuration and the change history that produced it.
Relationship to Periodic Review
Change control evaluates individual changes as they occur. Cleaning Validation Periodic Review and Lifecycle Assessment evaluates the accumulated evidence and determines whether the overall validation program still remains appropriate.
The periodic review should confirm that changes were assessed properly, required verification was completed, revalidation decisions remain justified, and cumulative small changes have not collectively moved the process beyond the state represented by the original validation.
Several individually minor changes can become significant when considered together.
This is one reason lifecycle assessment should not be replaced by individual change records alone.
Common Deficiencies
A common deficiency is defining a fixed rule that every cleaning-related change automatically requires full revalidation. This creates unnecessary work without improving scientific control and can obscure which validation assumptions actually changed.
The opposite deficiency is more serious: approving changes because they are administratively classified as minor without assessing their effect on cleanability, residue limits, sampling, analytical capability, or equipment configuration.
Other weaknesses include introducing new products without reassessing worst-case grouping, failing to recalculate MACO after an HBEL or dose change, changing product-contact materials without assessing recovery, modifying rinse or extraction volume without recalculating analytical criteria, and treating software recipe changes solely as computerized-system changes even when they alter the physical cleaning process.
Further deficiencies include describing materially different replacement parts as like-for-like, extending dirty hold or campaign length without supporting evidence, changing detergent concentration without assessing cleaning efficacy, removing worst-case sampling locations because they historically passed, and assuming that a more sensitive analytical method automatically requires a lower cleaning limit.
A lifecycle program can also fail through cumulative-change blindness. Several small changes to products, equipment, cleaning parameters, and analytical methods may individually appear acceptable while collectively invalidating the original validation assumptions.
Key Principles
- Cleaning-validation change control should protect the validated state by determining whether a proposed or observed change affects the assumptions and evidence supporting cleaning validation.
- Not every change requires revalidation. Where no significant cleaning-validation impact exists and existing evidence remains applicable, documented assessment can be sufficient. ICH Q7 similarly states that revalidation is normally unnecessary when no significant changes have occurred and review confirms continued acceptable performance.
- New products should be evaluated against existing worst-case products for both patient-safety limits and cleaning difficulty. Product grouping should be revised when a new product creates a more challenging condition.
- HBEL, PDE, or ADE changes should be propagated through MACO, surface and sample limits, analytical capability, worst-case rankings, and historical validation data before determining whether revalidation is necessary.
- Equipment modifications should be evaluated for changes to product-contact geometry, surface area, material, cleaning coverage, drainage, accessibility, and sampling recovery. A true like-for-like replacement may require little additional validation, while a changed product-contact design can require targeted or broad revalidation.
- Cleaning chemistry and process-parameter changes should be compared with the mechanism and operating range represented by validation. Changes that move the process outside the validated range require additional evidence.
- Automated recipe and software changes should be assessed for both computerized-system impact and cleaning-process impact whenever they can alter cleaning sequence, setpoints, valve positions, alarms, dosing, flow, time, or other validated conditions.
- Sampling, recovery, and analytical changes can affect the measurement system without changing cleaning effectiveness. These changes frequently require targeted method or recovery work rather than repetition of the entire cleaning validation.
- Revalidation scope should match the actual impact: no additional activity, documented verification, targeted bridging, targeted revalidation, or broader revalidation.
- Change control should be completed with implementation verification and documentation updates so the current cleaning-validation package accurately represents the products, equipment, methods, limits, and cleaning process actually in use.