

Analytical Method Validation for Cleaning Residue Testing
Cleaning-validation decisions depend on analytical results that can reliably distinguish acceptable residue levels from unacceptable carryover. After an analytical technology has been selected and the required sensitivity has been defined, the complete analytical procedure must be demonstrated to perform appropriately for its intended cleaning-residue application.
Analytical method validation for cleaning residue testing extends beyond instrument response. Depending on the procedure, the validated method can include sample preparation, extraction, dilution, standards and reagents, analytical measurement, calculations, data processing, and reporting. Swab extracts and rinse samples can also introduce detergent residues, excipients, sampling-material extractables, rinse-water background, and other components that influence the analytical result.
The objective is therefore not merely to demonstrate that an HPLC system, TOC analyzer, UV-visible spectrophotometer, conductivity meter, or other instrument functions correctly. Instrument qualification supports measurement reliability; analytical procedure validation demonstrates that the complete method is fit to make the intended cleaning decision. Selection of the analytical technology itself is addressed in Analytical Method Selection for Cleaning Validation.
Regulatory and Scientific Basis
21 CFR 211.160 — General Requirements requires scientifically sound specifications, standards, sampling plans, and test procedures as part of laboratory controls. It also requires laboratory instruments to be calibrated under established written programs.
21 CFR 211.194 — Laboratory Records requires laboratory records to identify the methods used and the location of data demonstrating that those methods meet appropriate standards of accuracy and reliability. It also requires suitability of testing methods to be verified under actual conditions of use and requires documentation supporting modifications to established methods.
Cleaning itself is governed by 21 CFR 211.67 — Equipment Cleaning and Maintenance. Analytical methods provide part of the evidence demonstrating that the approved cleaning process achieves the established residue requirements.
FDA’s Guide to Inspections: Validation of Cleaning Processes specifically addresses analytical-method sensitivity and the need to understand sampling recovery. The analytical and sampling procedures therefore need to function as parts of one cleaning-validation measurement system.
The current analytical framework is provided by FDA’s final ICH Q2(R2) Validation of Analytical Procedures and ICH Q14 Analytical Procedure Development guidances. Q2(R2) establishes a general framework for demonstrating analytical procedure performance, while Q14 addresses science- and risk-based analytical procedure development and lifecycle management.
Intended Use Defines the Validation Strategy
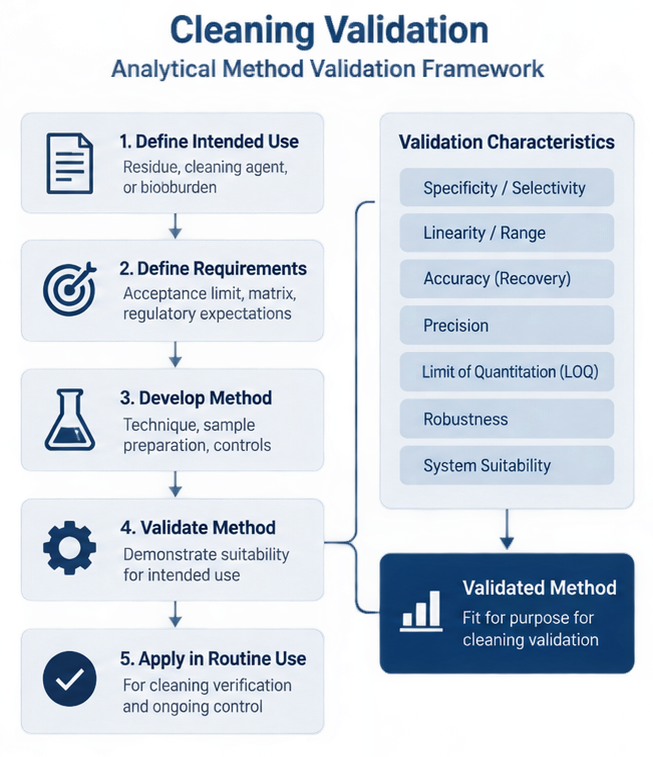
The validation strategy should begin with a clearly defined analytical intended use. The method should identify what residue or residue class is being measured, the sample type, applicable acceptance criterion, reportable units, expected concentration range, analytical technology, and the decision that will be made from the result.
A procedure intended to quantify a specific active pharmaceutical ingredient in a swab extract requires a different validation emphasis from a nonspecific TOC procedure used to conservatively represent organic residue. Likewise, conductivity used to evaluate an ionic cleaning agent or rinse endpoint requires controls different from those needed for chromatographic quantitation.
The analytical acceptance requirement should be linked to Cleaning Validation Acceptance Criteria and Surface, Swab, and Rinse Limits, while the required lower-range sensitivity should be established through Analytical Sensitivity and Quantitation Limits in Cleaning Validation. Validation should challenge the procedure in the concentration region where the cleaning decision will actually be made rather than only at analytically convenient concentrations.
Validate the Complete Analytical Procedure
The validated boundary should represent the procedure as it will actually be performed. Validation limited to detector response or calibration standards may underestimate variability introduced by sample preparation, extraction, dilution, filtration, reagent preparation, analyst technique, integration, calculations, and data processing.
This distinction is particularly important in cleaning validation because analyte concentrations are often low and preparation steps can materially affect the final result. A procedure that performs well using standards prepared directly in mobile phase may behave differently when applied to an extracted swab containing sampling-material extractables, residual detergent, excipients, or other matrix components.
ICH Q2(R2) specifically places accuracy assessment under normal analytical conditions, including relevant sample matrix and the described sample-preparation procedure. The validation study should therefore represent the approved extraction solvent, extraction volume, dilution sequence, containers, holding conditions, filtration where applicable, instrument conditions, calculations, correction factors, and reporting convention.
Specificity and Selectivity
Specificity and selectivity establish whether the analytical procedure can determine the residue of interest without unacceptable influence from other materials likely to be present. ICH Q2(R2) recognizes demonstration through absence of interference, comparison with an orthogonal analytical procedure, or—in justified circumstances—the inherent scientific characteristics of the measurement technology.
For cleaning-residue testing, potential interferences extend beyond those encountered in routine finished-product analysis. They may include excipients, other product residues, degradation products, detergents, sanitizers, process materials, swab extractables, extraction solvents, rinse-water background, containers, and substances introduced during sample preparation.
For HPLC or UHPLC procedures, the target analyte should be adequately discriminated from relevant matrix components and other chromatographic responses. Where degradation during manufacturing, dirty hold time, cleaning, sample storage, or extraction is credible, the potential effect of degradation products should be considered.
UV-visible methods require particular attention because spectral response may be shared by the analyte, excipients, detergents, or degradation products. Absence of chromatographic separation can make an apparently sensitive method unsuitable when the measured absorbance cannot be attributed reliably to the target residue.
TOC and conductivity require a different interpretation. These technologies are intentionally nonspecific. Their validation strategy should demonstrate that background is adequately controlled and that the aggregate response provides scientifically justified evidence for the residue-control objective. Lack of compound specificity does not make a method invalid, but it changes what the result can legitimately demonstrate.
Sample Matrix and Interference Challenges
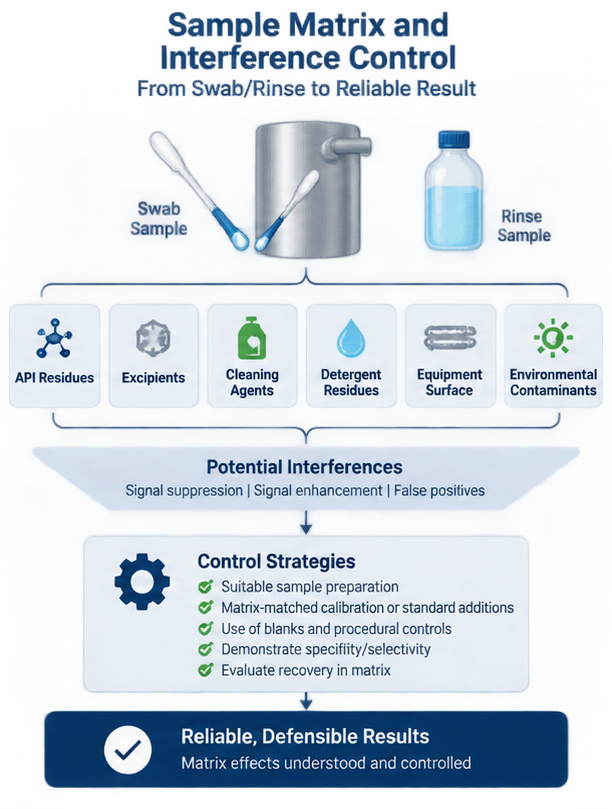
Cleaning-validation methods should be challenged using matrices representative of routine samples where the matrix can influence analytical performance. A standard prepared in clean solvent does not necessarily reproduce the conditions encountered in a swab extract or equipment rinse.
Representative challenges may include extracted unused swabs, rinse water, extraction solvent, detergent, placebo or excipient components, degradation products, and other materials identified through process knowledge as credible sources of interference. The objective is not to combine every conceivable substance into one artificial mixture. The study should represent materials that could realistically alter the analytical result.
Matrix effects can produce response enhancement or suppression, interfere with analyte identification, alter extraction performance, increase baseline variability, or generate positive bias. These effects are particularly significant near the lower end of the analytical range because a small background contribution can represent a substantial portion of the reported residue concentration.
Blanks and Background Control
Blank response should be understood as part of low-level method capability. An instrument blank or solvent blank evaluates only a portion of the analytical process. A procedural blank that includes sampling material, solvent, container, extraction, dilution, and other preparation steps can provide a more representative assessment of background contribution.
For swab methods, background can originate from the swab itself, extraction solvent, containers, filters, or handling. Rinse methods can be affected by background in the rinse medium and collection system. TOC methods require particular control because organic material from water, containers, swabs, or handling contributes directly to the measured carbon response.
Blank subtraction should not be used automatically to manufacture an apparently lower quantitative limit. If blank correction is part of the approved analytical procedure, background variability, calculation method, and impact on low-level accuracy should be established during development and validation.
Accuracy
Accuracy describes the closeness of the measured result to an accepted or known value. ICH Q2(R2) states that accuracy should be established across the reportable range and under the normal test conditions of the procedure, including relevant sample matrix and sample preparation.
For cleaning-residue methods, accuracy is commonly evaluated using samples fortified with known quantities of analyte at concentrations spanning the intended range. The design should include the concentration region associated with the cleaning acceptance criterion, because low-level bias may not be apparent from results generated only at higher concentrations.
Accuracy may be expressed as percent recovery of a known added amount or through another justified comparison with an accepted value. The number of concentration levels and replicates should be appropriate to the procedure, intended range, analytical risk, and available prior knowledge rather than copied mechanically from another validation protocol.
Analytical accuracy must be distinguished from surface sampling recovery. Analytical accuracy determines whether the procedure measures the analyte present in the analytical sample correctly. Surface recovery evaluates how effectively the swab or rinse procedure removes residue from the equipment surface and transfers it into the sample. These are related but different validation questions.
Precision
Precision describes agreement among repeated measurements. Cleaning-residue methods should evaluate precision under conditions representative of actual use rather than relying solely on replicate injections from one prepared standard.
Repeatability evaluates variation under the same operating conditions over a short interval. Intermediate precision evaluates relevant within-laboratory variables such as analysts, days, instruments, reagent preparations, columns, or other conditions expected during routine application. Reproducibility may become relevant where the method is transferred or routinely performed at multiple laboratories.
ICH Q2(R2) recognizes repeatability, intermediate precision, and reproducibility as distinct levels of precision and recommends statistical characterization of the observed variability.
Performance near the cleaning acceptance criterion deserves particular attention. A method may show excellent precision at moderate concentrations while becoming highly variable near its lower quantitative range. Validation should therefore demonstrate adequate precision where the cleaning decision is made.
Response, Linearity, and Calibration Model
Where the analytical procedure is expected to generate a linear relationship between analyte concentration and measured response, that relationship should be demonstrated across the intended range. Evaluation should consider the slope, intercept, regression characteristics, and distribution of residual error rather than relying only on a high correlation coefficient.
ICH Q2(R2) recommends a minimum of five appropriately distributed concentrations when linearity is assessed and specifically notes the value of evaluating deviations from the regression line. It also recognizes analytical procedures that legitimately use nonlinear or multivariate calibration models.
For cleaning-residue methods, the calibration or working range should represent concentrations expected after sampling, extraction, recovery, dilution, and other preparation steps. An unnecessarily broad calibration range can obscure performance in the low concentration region that actually governs the cleaning decision.
A high coefficient of determination does not by itself establish quantitative method capability. The calibration model must work together with acceptable accuracy and precision across the relevant range.
Range and Quantitative Method Capability
Quantitative method capability is not an independent validation characteristic. It is the combined ability of the analytical procedure to generate reliable numerical results across the concentration interval required for its intended use.
ICH Q2(R2) defines analytical range in terms of an interval over which the procedure provides suitable response, accuracy, and precision. It also distinguishes the concentration range presented to the instrument from the reportable range used for the final analytical result.
For cleaning validation, the applicable cleaning acceptance criterion should fall within the demonstrated quantitative capability of the procedure. This relationship should be evaluated after considering sampled area, extraction volume, rinse volume, dilution, recovery treatment, and calculation factors used to convert the analytical result into the units used for the cleaning decision.
The lower end of the range is closely related to the method’s limit of quantitation. Analytical Sensitivity and Quantitation Limits in Cleaning Validation addresses LOD, LOQ, background effects, lower-range verification, and conversion of analytical sensitivity into practical surface or rinse capability.
A numerical instrument output should not automatically be treated as a quantitatively valid result. The reported concentration should fall within a region where the complete procedure has demonstrated acceptable performance.
Extraction Efficiency and Sampling Recovery
Cleaning-residue testing can involve two different recovery mechanisms that should not be confused. First, the sampling procedure must recover residue from the equipment surface. Second, the analytical preparation may need to extract that recovered residue from the swab or other sampling device into a solution suitable for measurement.
The complete relationship can be viewed as: Residue on equipment → surface sampling recovery → extraction into analytical sample → analytical measurement → calculated cleaning result
The first component is addressed in Swab and Rinse Recovery Studies for Cleaning Validation. The second can be part of the analytical procedure itself and should be evaluated during method development and validation where it can materially affect the result.
Extraction solvent, extraction time, agitation or sonication, temperature, sample volume, holding time, and other preparation variables may affect recovery. Low but reproducible extraction can sometimes be managed through a scientifically justified procedure, but poor or highly variable extraction reduces confidence in results near the acceptance criterion.
The approved procedure should also define how surface recovery is treated in calculations. Some programs apply a validated recovery correction factor, while others demonstrate adequate recovery but intentionally report uncorrected results as part of a conservative strategy. Whichever approach is selected should be documented and applied consistently.
Detergent and Excipient Interference
Cleaning-agent and excipient interference deserves explicit evaluation because these materials can remain at low concentrations and can affect different analytical technologies in different ways.
In chromatographic procedures, detergent components or excipients can co-elute, alter peak shape, affect retention, or interfere with detector response. In UV-visible methods they may contribute directly to absorbance. Organic detergents contribute to TOC and cannot be distinguished from other organic residues by TOC alone. Conductivity responds to ionic species regardless of their source.
The study design should therefore challenge credible materials at concentrations appropriate to the cleaning application. Where one analytical procedure cannot provide adequate discrimination, use of an additional or orthogonal procedure may be justified. ICH Q2(R2) specifically recognizes combination of procedures when one procedure alone does not provide sufficient specificity or selectivity.
Robustness
Robustness evaluates whether the analytical procedure continues to meet expected performance when normal variations occur in relevant parameters. ICH Q14 places robustness primarily within analytical procedure development, while Q2(R2) recognizes it as a measure of the procedure’s capacity to meet expected performance criteria during normal use.
Potential variables for cleaning-residue methods can include extraction time, extraction solvent composition, pH, mobile-phase preparation, column temperature, flow rate, wavelength, instrument settings, reagent lot, sample holding time, analyst handling, and other parameters identified during method development.
Not every parameter needs to be studied with equal intensity. The robustness strategy should focus on variables that can realistically affect results near the cleaning acceptance criterion.
Sample and standard stability should be included where holding time could alter the result. This can be especially important for low-level residues, unstable compounds, TOC samples susceptible to external organic contamination, or samples where adsorption to the container can occur.
Suitability at the Cleaning Acceptance Limit
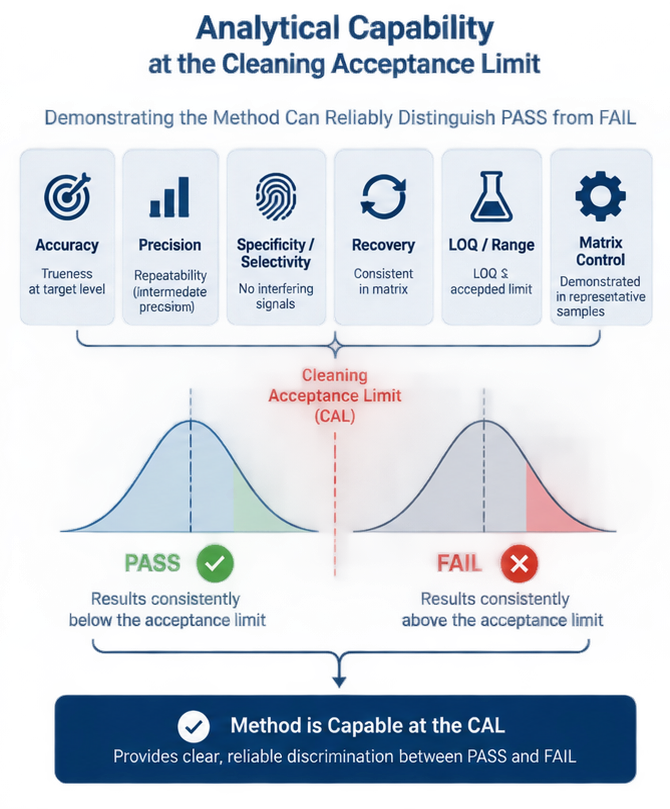
The most important validation question is whether the analytical procedure can make the intended cleaning decision reliably. Passing generic accuracy, precision, or linearity requirements at concentrations unrelated to the cleaning limit does not answer that question.
The method should demonstrate suitable performance at and around the concentration corresponding to the cleaning acceptance criterion. Accuracy, precision, specificity or selectivity, recovery, matrix effects, blank response, LOQ, and validated range collectively determine whether the analytical result can discriminate an acceptable cleaning condition from an unacceptable one.
The acceptance criterion should be traced through the complete measurement chain: Cleaning surface or equipment limit → sampled residue amount → extraction or rinse concentration → analytical working concentration → measured result → conversion back to cleaning acceptance units
Each transformation should be understood and controlled. Quantitative results should not depend on unjustified extrapolation below the validated range or on assumptions about sampling recovery or matrix behavior that were not demonstrated experimentally.
Technique-Specific Validation Considerations
The relative importance and manner of demonstrating validation characteristics depend on the analytical technology. HPLC and UHPLC methods generally require strong evaluation of chromatographic discrimination, calibration response, lower-range capability, sample preparation, and potential matrix peaks. The associated analytical system should also remain appropriately qualified and controlled for its GMP intended use.
UV-visible methods can provide efficient quantitative testing when absorbance characteristics and matrix composition support adequate selectivity. Because the technique lacks chromatographic separation, interference and background response can become primary limitations.
TOC methods require demonstration that the residue contributes measurable oxidizable carbon under the selected conditions and that background carbon is sufficiently controlled. Where TOC is used conservatively, the rationale for assigning measured carbon to the residue or residue group should be documented.
Conductivity methods should demonstrate a meaningful relationship between ionic residue concentration and measured conductivity under the relevant water quality and temperature conditions. Background conductivity and unrelated ionic species must not prevent reliable interpretation at the required cleaning limit.
The analytical technology should therefore determine the validation design rather than forcing every method through an identical checklist.
Validation Protocol and Acceptance Criteria
The validation protocol should define the analytical intended use, method boundary, responsibilities, sample types, validation characteristics, experimental design, concentration levels, replicates, matrices, predefined acceptance criteria, calculations, statistical evaluations, deviation handling, and final approval requirements.
Acceptance criteria should be justified from method development knowledge, intended use, analytical technology, concentration range, and the risk associated with measurement error. Generic criteria copied from another procedure can be misleading where residue concentrations, sample matrices, or analytical principles differ.
Validation deviations should be assessed for their potential impact on the study conclusion. Failure of a predefined validation criterion should be investigated before the procedure is approved for cleaning-validation use.
The final validation report should summarize execution, results, statistical evaluations, deviations, investigations, approved range, method limitations, and the conclusion that the procedure is suitable for the intended cleaning-residue application.
System Suitability Is Not Method Validation
Analytical method validation and routine system suitability serve different purposes. Validation establishes that the analytical procedure can perform appropriately for its intended use. System suitability provides evidence that the analytical system is functioning acceptably when samples are tested.
For chromatographic methods, system suitability may address response precision, retention, resolution, peak shape, or other method-critical characteristics. Other technologies require controls appropriate to their measurement principles.
Passing system suitability does not demonstrate that the method is appropriate for the cleaning sample matrix or residue limit. Conversely, a validated method should not be used when the analytical system fails its approved routine suitability criteria.
Analytical Method Transfer
When a validated cleaning-residue procedure is transferred to another laboratory, the transfer should demonstrate that the receiving laboratory can execute the procedure reliably under its own operating conditions.
Transfer activities may include comparative testing, precision studies, recovery evaluation, system-suitability confirmation, and review of critical sample-preparation steps. The extent should reflect method complexity, equipment equivalence, analyst experience, sample matrix, and intended use.
Full revalidation is not automatically necessary solely because a method is transferred. Significant changes in equipment, procedure, matrix, preparation, or operating conditions may, however, require targeted or broader revalidation.
Lifecycle Management and Change Control
Analytical procedure validation is not a one-time activity. ICH Q14 establishes a lifecycle approach in which knowledge generated during development and validation supports control of the analytical procedure and scientifically justified change management.
Changes should be assessed according to their potential effect on validated performance and the cleaning decision. Relevant changes can include tighter residue limits, new products or cleaning agents, different swab materials, revised extraction solvents or volumes, altered dilution schemes, new matrix components, changes in chromatographic columns or conditions, detector replacement, revised software calculations, sample holding-time changes, or method transfer.
The impact assessment should determine whether the change requires documentation only, targeted verification, partial revalidation, or broader revalidation. Q2(R2) recognizes that revalidation may address all or only selected analytical performance characteristics depending on the nature of the change.
Method performance should also be reconsidered when routine experience identifies increasing blank response, unusual variability, declining recovery, repeated system-suitability failures, atypical results, investigation trends, or other evidence that the analytical procedure may no longer perform as originally demonstrated.
Relationship to the Cleaning Validation Measurement System
Analytical method validation is only one component of the complete cleaning-validation evidence chain. A strong analytical method cannot compensate for poorly selected sampling locations, inadequate surface recovery, an inappropriate residue limit, or a cleaning process that lacks reproducibility.
Similarly, an effective cleaning process cannot be convincingly demonstrated when the analytical method cannot reliably measure the required residue level. Cleaning validation therefore depends on alignment among the cleaning process, acceptance limit, sampling location, swab or rinse technique, sampling recovery, analytical procedure, and final calculation.
Cleaning Validation Sampling Strategy and Worst-Case Locations establishes where evidence should be collected. Swab Sampling for Cleaning Validation and Rinse Sampling for Cleaning Validation address how the samples are obtained. Swab and Rinse Recovery Studies for Cleaning Validation establishes recovery capability. Analytical method validation then demonstrates that those samples can be measured with sufficient reliability to support the cleaning decision.
Common Validation Deficiencies
Common deficiencies include validating only standard solutions instead of representative sample matrices; evaluating instrument response while excluding extraction or sample-preparation steps; relying on a high correlation coefficient as the principal evidence of quantitative capability; failing to challenge detergent, excipient, swab, or rinse-water interference; and validating a range that does not adequately include the cleaning acceptance criterion.
Other weaknesses include confusing analytical accuracy with surface recovery, applying generic validation criteria without scientific justification, using a nonspecific method without defining how the aggregate response will be interpreted, failing to understand blank contribution at low concentrations, reporting numerical values outside the demonstrated quantitative range, and assuming that instrument qualification eliminates the need for analytical procedure validation.
Lifecycle deficiencies include modifying extraction conditions, sample preparation, chromatographic conditions, software calculations, sample holding times, or other critical procedure elements without evaluating whether the original validation remains applicable.
Key Principles
Analytical method validation for cleaning residue testing should demonstrate that the complete analytical procedure is fit for the intended cleaning decision. The validation strategy should reflect the residue, analytical technology, sample matrix, applicable acceptance criterion, and intended use rather than applying the same test checklist to every method.
Specificity or selectivity, accuracy, precision, response, range, lower-range capability, extraction, matrix effects, blanks, and robustness should be evaluated where relevant. Cleaning-residue applications require particular attention to detergents, excipients, sampling materials, rinse-water background, recovery, and other effects that may not be represented by conventional analytical standards.
Quantitative method capability is the combined result of these characteristics. It is demonstrated when the complete procedure can generate reliable numerical results at and around the cleaning acceptance criterion.
Instrument qualification, analytical method validation, sampling recovery, and system suitability provide different forms of evidence and should not be treated as interchangeable. The analytical procedure should remain under lifecycle control, and changes that can affect its ability to support the cleaning decision should be assessed for verification or revalidation.