

Package Seal Strength, Integrity, and Sterile Barrier Testing
Packaging integrity testing provides evidence that a packaging system continues to perform the containment, barrier, and protective functions established during packaging design and qualification. For sterile medical-device packaging, this includes demonstrating that the sterile barrier system remains intact after packaging, sterilization, aging, handling, and distribution challenges. For pharmaceutical products, integrity can involve containment, protection from environmental exposure, or maintenance of a sterile container-closure system.
A sound test strategy begins by defining what characteristic is being evaluated. Seal strength, seal quality, leak detection, package integrity, and container-closure integrity answer different questions. A strong seal is not necessarily leak-tight. A visually acceptable seal may contain a channel. A package that passes an integrity test may still have inadequate opening characteristics. Conversely, a cosmetic defect does not necessarily constitute loss of package integrity.
For terminally sterilized medical devices, ISO 11607-1:2019 establishes requirements for materials, sterile barrier systems, and packaging systems and remains current following systematic review; Amendment 1:2023 adds application of risk management. FDA’s current recognized-consensus-standards database also includes multiple ASTM methods used to evaluate medical packaging, including ASTM F88/F88M-23 for seal strength and ASTM F1929-23 for dye-penetration testing of porous medical packaging.
For sterile pharmaceutical and biological products, package testing should remain aligned with the separate concept of container-closure integrity testing (CCIT). FDA’s final guidance recognizes validated physical, chemical, and microbiological integrity methods for demonstrating continued container-closure integrity during shelf life, while making clear that integrity testing does not replace initial product sterility testing. Detailed pharmaceutical CCIT strategy is addressed separately in Container Closure Integrity Testing (CCIT).
Key Principles
- Seal strength, seal quality, and package integrity should be treated as different package characteristics.
- No single test method demonstrates every aspect of package integrity.
- Test selection should be based on package materials, barrier function, anticipated defect type, required sensitivity, and intended use.
- Visual inspection is an important package-quality control but should not automatically be treated as proof of leak-tight integrity.
- Seal-strength testing measures mechanical separation force and does not directly demonstrate absence of channels or leaks.
- Sterile-barrier testing should evaluate the complete barrier, including seals, material surfaces, closures, and other critical interfaces.
- Packaging should be evaluated after the processing and lifecycle stresses relevant to the qualification claim.
- Worst-case configurations should be selected according to the failure mechanism being evaluated.
- Test methods should be demonstrated suitable for the specific package configuration and defect types they are expected to detect.
- Acceptance criteria should distinguish functional package failures from cosmetic observations that do not affect required package performance.
Distinguishing Seal Strength, Seal Quality, and Package Integrity
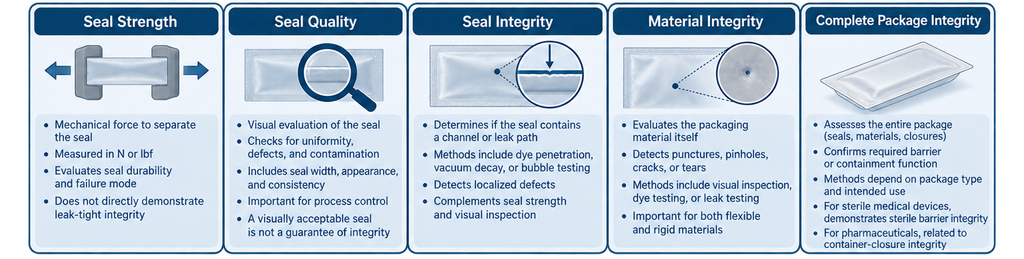
The term packaging integrity testing is often used too broadly. Several distinct characteristics should be separated before the test program is developed.
| Characteristic | Primary question |
|---|---|
| Seal strength | How much force is required to separate or fail the seal? |
| Seal quality | Was the seal formed uniformly and without observable defects? |
| Seal integrity | Does the sealed interface contain a channel, gap, or other leak path? |
| Material integrity | Does the package substrate contain punctures, cracks, pinholes, or other breaches? |
| Package integrity | Does the complete package maintain its required barrier or containment function? |
| Sterile barrier integrity | Does the medical-device sterile barrier remain capable of maintaining the required microbial barrier? |
| Container-closure integrity | Can the pharmaceutical container-closure system maintain the required barrier against ingress or egress? |
These characteristics overlap but cannot substitute automatically for one another.
For example, ASTM F88/F88M measures the force required to separate a seal and identifies the mode of failure. ASTM describes seal strength as useful for process validation, capability, and control, but this measurement is fundamentally a mechanical property. A pouch can have high seal strength and still contain a narrow channel created by contamination or local incomplete sealing.
Likewise, an acceptable dye-penetration test can provide evidence that a specified type of channel was not detected, but it does not quantify the mechanical strength of the seal.
Sterile Barrier Systems
For terminally sterilized medical devices, the sterile barrier system is the minimum packaging necessary to maintain the microbial barrier and permit aseptic presentation. Protective packaging outside that barrier may be necessary to prevent physical damage during storage and distribution. These relationships are discussed in Packaging System Types and Sterile Barrier Systems.
ISO 11607-1 addresses the materials, sterile barrier systems, and packaging systems used for terminally sterilized medical devices. ISO 11607-2 separately addresses validation of the forming, sealing, and assembly processes used to produce those systems. Both 2019 editions remain published and were confirmed through ISO’s 2024 review process; both also have 2023 amendments addressing risk management.
The sterile-barrier assessment should therefore consider more than the seal line. Potential integrity failures can occur through:
- seal channels or incomplete seals;
- seal separation;
- punctures;
- abrasion;
- flex cracking;
- tears;
- material pinholes;
- tray or lid damage;
- delamination;
- folds or wrinkles entering a seal;
- foreign material within the seal;
- damage caused by the device itself.
The test strategy should reflect the credible failure mechanisms identified during packaging risk assessment.
Pharmaceutical Container-Closure Integrity
Pharmaceutical container-closure systems require a different regulatory and technical framework from medical-device sterile-barrier packaging.
21 CFR §211.94 requires drug-product containers and closures to provide adequate protection against foreseeable external factors that can cause deterioration or contamination. FDA also permits appropriately validated container-closure integrity methods to support continued sterility assessment within stability programs for sterile products.
FDA specifically notes that acceptable integrity approaches can include pressure or vacuum decay, trace-gas methods, dye penetration, seal-force measurements, electrical methods, and microbiological challenge methods when properly validated for the particular container-closure system. FDA does not prescribe one universal integrity method or acceptance limit.
USP General Chapter <1207> and its related chapters provide the compendial framework for package integrity evaluation of sterile pharmaceutical products. Because this website has a dedicated Container Closure Integrity Testing (CCIT) article, detailed treatment of deterministic versus probabilistic CCIT technologies should remain there rather than being duplicated in this packaging article.
The distinction is important: ISO 11607 sterile-barrier testing should not be automatically applied to pharmaceutical vials, syringes, cartridges, or ampoules as if they were medical-device pouches or trays.
Visual Inspection
Visual inspection is often the first integrity-related assessment because many package failures produce visible evidence.
Depending on the package, inspectors may look for:
- incomplete seals;
- wrinkles or folds crossing the seal;
- contamination within the seal;
- delamination;
- tears;
- punctures;
- material thinning;
- tray cracks;
- distorted components;
- seal-width irregularities;
- peeling or separation;
- abrasion damage.
ASTM F1886/F1886M-25 provides a current method for visual evaluation of seals and packaging-material surfaces. The standard emphasizes that visual findings can indicate potential integrity defects but that additional testing may be required to determine whether an observation represents an actual integrity breach.
This is the proper role of visual inspection. It is valuable for detecting manufacturing variation and physical damage, but absence of a visible defect should not automatically be interpreted as proof of package integrity.
Where visual inspection is used as a controlled test method, lighting, viewing conditions, inspector training, defect standards, package orientation, and defined defect classifications should be established sufficiently to support consistent interpretation.
Seal Strength Testing
Seal-strength testing evaluates the mechanical force required to separate a sealed interface.
ASTM F88/F88M-23 is the principal ASTM method for measuring seal strength of flexible barrier materials. It can be applied to flexible-to-flexible and flexible-to-rigid or semi-rigid seals and provides information about both separation force and specimen failure mode. ASTM identifies seal strength as useful in packaging-process validation, process capability, and control. FDA currently recognizes ASTM F88/F88M-23 as a consensus standard within its medical-device sterility category.
Seal-strength evaluation can support:
- establishment of minimum seal performance;
- packaging-process development;
- comparison of sealing conditions;
- process capability assessment;
- aging studies;
- post-sterilization evaluation;
- post-distribution evaluation;
- investigation of package failures.
However, seal strength must not be described as equivalent to seal integrity.
A package with acceptable average seal strength can contain a localized channel. Likewise, a package with an extremely strong seal can create other problems, including material tearing or excessive opening force. The appropriate acceptance range should therefore reflect both package protection and intended use.
For peel-open medical-device packages, opening performance and aseptic presentation can also be relevant. The objective is not necessarily to produce the strongest possible seal, but to establish a reproducible seal strong enough to maintain package integrity while remaining appropriate for intended opening and use.
Burst and Pressure-Based Seal Strength Tests
Internal pressurization provides another way to characterize package mechanical performance.
ASTM F2054/F2054M describes burst testing of flexible package seals using internal air pressurization within restraining plates. The method increases package pressure until seal failure occurs and can help identify the weakest area of the seal. ASTM specifically cautions that burst-test results do not necessarily correlate directly with seal strength measured using ASTM F88 or similar methods.
This reinforces an important validation principle: different mechanical tests measure different package behaviors.
Burst testing may be useful when the qualification objective concerns resistance to pressure differential, but it should not automatically be presented as a leak-detection method or as a replacement for localized seal-strength testing.
Dye-Penetration Testing
Dye penetration is commonly used to identify channel-type defects in package seals.
ASTM F1929-23 applies to seals formed between a transparent material and a porous sheet material. The current standard describes procedures capable of detecting and locating channels represented by a 50 µm wire under the defined test conditions. FDA currently recognizes F1929-23 for medical-device packaging applications.
For nonporous packaging and flexible barrier materials, ASTM F3039-23 provides a corresponding dye-penetration approach. ASTM describes this as a destructive, qualitative go/no-go method rather than a quantitative measurement of leak size. FDA also lists F3039-23 among its recognized medical-device sterility standards.
Dye testing is useful because it can help locate a seal channel. Its limitations should also be understood:
- the test is destructive;
- interpretation can depend on visual observation;
- sensitivity depends on the package and method conditions;
- it does not measure seal strength;
- it does not provide a quantitative leak rate;
- applicability depends on the package materials.
Accordingly, selection of dye penetration should be justified by the defect mechanism and package construction rather than simply because it is historically familiar.
Bubble Leak Testing
ASTM F2096 provides a gross-leak test using internal pressurization while the package is submerged. Escaping bubbles indicate a leak path.
The current ASTM listing describes the method as applicable to tray and pouch packages and states a demonstrated sensitivity down to approximately 250 µm under the specific conditions used for the method’s evaluation. The test is destructive because the package must be entered to supply internal pressure.
Bubble testing can be useful for:
- locating relatively large leaks;
- investigating damaged packages;
- development work;
- examining packages that are difficult to test by other approaches.
It should not be represented as a highly sensitive universal integrity test. A package can pass a gross-leak test while still containing a smaller defect relevant to another product or barrier requirement.
Vacuum Decay Testing
Vacuum decay provides an instrumental approach to package leak detection. The package is placed in an evacuated chamber, and the system measures changes associated with gas or vapor escaping through a leak.
ASTM F2338-24 covers nondestructive vacuum-decay testing for several rigid, semi-rigid, flexible, porous-lidded, and nonporous package configurations. The actual sensitivity depends on the product-package system and test configuration and therefore must be demonstrated for the specific application.
Vacuum decay can offer advantages where an objective instrumental response, nondestructive testing, or increased repeatability is required. However, the presence of an ASTM method does not eliminate the need to establish appropriate fixtures, test parameters, detection capability, positive controls, and package-specific acceptance criteria.
Selecting the Test Method
Method selection should begin with the required packaging function and expected defect mechanism.
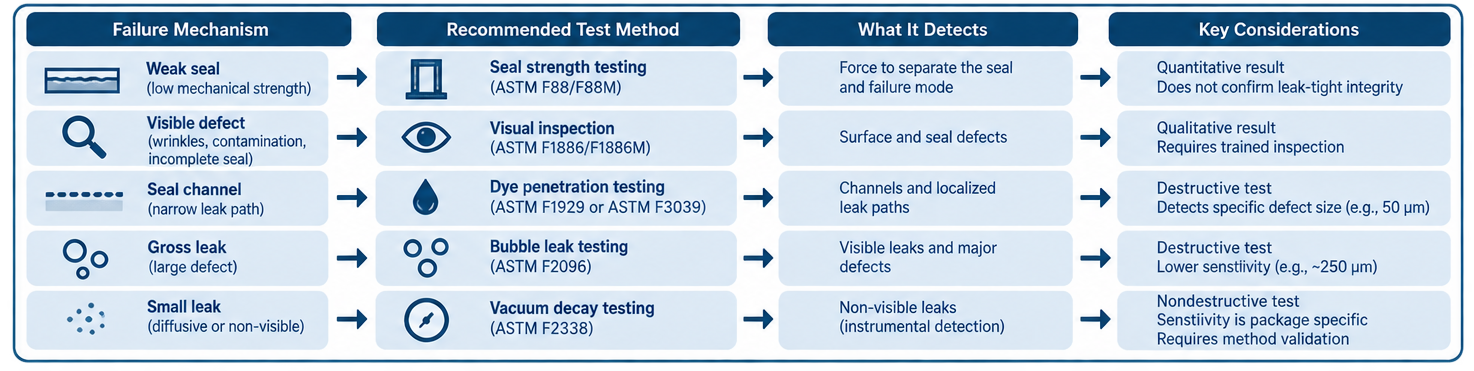
A practical decision sequence is: Required barrier function → credible defect → package material/configuration → required detection capability → suitable test technology → validated method → acceptance criterion
The following comparison illustrates why no method should be selected in isolation.
| Method | Primary information | Typical output | Key limitation |
|---|---|---|---|
| Visual inspection | Visible seal/material defects | Qualitative | Cannot demonstrate absence of all leak paths |
| ASTM F88/F88M | Seal separation force | Quantitative force | Does not directly measure leakage |
| ASTM F2054/F2054M | Burst resistance | Quantitative pressure | Not equivalent to local seal strength or leak testing |
| ASTM F1929 | Channels in porous-package seals | Pass/fail + defect location | Destructive and qualitative |
| ASTM F3039 | Leaks in nonporous package seals/materials | Pass/fail + defect location | Destructive and qualitative |
| ASTM F2096 | Gross package leaks | Visual bubble indication | Relatively coarse leak detection |
| ASTM F2338 | Package leakage by vacuum decay | Instrumental response | Package-specific setup and sensitivity required |
The correct strategy frequently uses complementary methods rather than searching for one test to answer every question.
Test Method Suitability and Validation
A standard test designation does not by itself demonstrate that the method is suitable for a specific product-package configuration.
Method validation or method-suitability work should establish, as applicable:
- package types and materials covered;
- defect types the method is intended to detect;
- applicable detection capability;
- positive and negative controls;
- test equipment and fixtures;
- sample preparation;
- environmental conditions;
- repeatability;
- operator effects where interpretation is involved;
- acceptance criteria;
- known limitations.
For instrumental leak methods, method development should establish discrimination between acceptable units and defined defective units. For visual or dye-based methods, controlled defect samples can be particularly useful for demonstrating inspector or method capability.
Acceptance limits should not be copied from another product merely because the same ASTM method is used. The relevant defect and required performance depend on the specific packaging system and its intended barrier function.
Positive Controls and Known Defects
Integrity-method qualification should include appropriately characterized defective units when technically feasible.
Known defects can help establish whether the selected method can detect the package failures important to the application. Depending on the technology, challenge units might represent:
- seal channels;
- pinholes;
- punctures;
- incomplete seals;
- material cuts;
- closure defects;
- controlled leak paths.
The artificial defect should represent the intended failure mechanism sufficiently to support the method claim. Randomly damaged packages may be useful during exploratory development but are less suitable when a defined detection capability must be demonstrated.
Positive controls also help distinguish method failure from package performance. If a challenge defect is not detected, the issue may lie with the method or execution rather than with the package population being tested.
Sample and Worst-Case Selection
Sample selection should follow the same technical logic established in Packaging Qualification Strategy and Worst-Case Configuration. There is no universal worst case for integrity testing.
For seal testing, important variables can include:
- seal length and width;
- material combination;
- package dimensions;
- product load near the seal;
- sealing geometry;
- sterilization exposure;
- aging condition;
- device movement;
- product mass;
- material thickness.
The configuration representing maximum drop impact may not represent the worst seal-integrity condition. The test protocol should therefore identify why the selected units challenge the particular characteristic being measured.
When Integrity Testing Should Be Performed
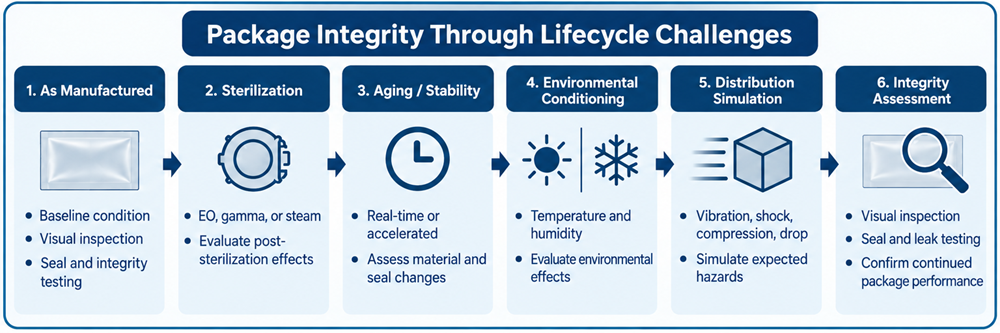
Integrity should be evaluated at lifecycle points relevant to the packaging claim.
Potential conditions include:
- As manufactured. Establishes baseline package condition and can help separate process-related defects from damage introduced later.
- After sterilization. Sterilization can alter films, adhesives, seals, trays, closures, and other materials.
- After aging. Material and seal properties can change during shelf life.
- After environmental conditioning. Temperature and humidity can affect package materials and seals.
- After distribution simulation. Vibration, shock, compression, drop, and abrasion can create package damage that is not present initially.
- After actual shipping, where appropriate. Real-world distribution studies can provide additional evidence concerning package performance within the commercial logistics system.
Not every product requires every condition as a separate study. The qualification strategy should determine which sequence provides the evidence required for the specific package and claim.
Relationship to Distribution Simulation
Package integrity testing is often most valuable when performed after controlled mechanical and environmental challenge.
Distribution Simulation Strategy and Transport Testing addresses selection and execution of vibration, shock, compression, drop, and other distribution challenges. Integrity testing then determines whether those challenges produced damage relevant to the required package function.
The relationship can be represented as: Qualified package configuration → conditioning → distribution challenge → visual inspection → integrity / seal / product evaluation
This sequence is particularly important for sterile medical-device packaging because damage may be caused by interaction between the device and sterile barrier during transportation. Abrasion, movement, concentrated loading, and impact can compromise the barrier even when the initial sealing process was fully capable.
Acceptance Criteria
Acceptance criteria should correspond to the characteristic being tested.
Examples include:
Seal-strength criteria
- minimum or defined range of separation force;
- acceptable failure mode;
- no unacceptable material tearing where peelability is required.
Visual criteria
- no channels of defined significance;
- no punctures or tears;
- no unacceptable seal folds or contamination;
- no material damage that compromises required package function.
Integrity criteria
- no detected leak above the validated method threshold;
- no dye penetration meeting the defined failure condition;
- no bubble emission meeting the defined defect criterion;
- instrumental response within the validated acceptance range.
The protocol should avoid a generic criterion such as “package intact.” The actual acceptance condition should be stated in terms appropriate to the method and required package function.
Integrity Failures and Investigation
An integrity failure should be investigated as a packaging-system failure until evidence establishes another cause.
Potential sources include:
- material defects;
- sealing-process variation;
- incorrect equipment settings;
- contamination within a seal;
- component dimensional variation;
- sterilization damage;
- aging degradation;
- device movement;
- abrasion;
- distribution stress;
- test-method or fixture problems;
- operator execution.
The investigation should establish whether the observed defect is isolated, systemic, process-related, design-related, or caused by the qualification challenge.
Repeating the test with new samples does not invalidate the original failure. Retesting should follow a documented investigation and scientifically justified disposition.
Connection to Packaging Process Validation
Integrity testing and packaging-process validation support one another but answer different questions.
Integrity testing asks whether the finished package performs its required barrier or containment function.
Packaging-process validation asks whether forming, sealing, and assembly operations consistently produce packages meeting defined requirements.
ISO 11607-2 specifically addresses development and validation of forming, sealing, and assembly processes for terminally sterilized medical-device packaging. The planned article Packaging Process Validation: Forming, Sealing, and Assembly will address that process-validation framework in detail.
Seal strength and integrity data can provide critical process evidence, but end-product testing should not be used to compensate for an inadequately controlled sealing process.
Lifecycle Control
Packaging integrity assumptions should be reassessed when changes can affect materials, seals, closures, package geometry, processing, or distribution stress.
Relevant changes can include:
- material or supplier changes;
- adhesive or coating changes;
- seal-width changes;
- package-size changes;
- new product or device configurations;
- sealing equipment or tooling changes;
- sterilization changes;
- extended shelf life;
- new distribution configurations;
- changes in protective packaging;
- new integrity-test equipment or methods.
The change assessment should determine whether existing qualification data remain representative or whether focused testing or requalification is required.
Validation Perspective
A defensible package integrity strategy should answer five separate questions:
- What package function must be maintained?
- What defect or failure mechanism could compromise that function?
- Which test method actually detects or measures that characteristic?
- Under what lifecycle conditions must the package continue to meet the requirement?
- What acceptance criterion distinguishes acceptable from unacceptable performance?
The most important validation distinction is that seal strength is not seal integrity, visual appearance is not proof of integrity, and package integrity is not automatically equivalent to pharmaceutical CCIT. Each provides different evidence. When the methods are selected according to function and failure mechanism, package testing becomes a coherent qualification program rather than a collection of familiar ASTM tests.