

Verification of CPPs and Process Control Strategy During PPQ
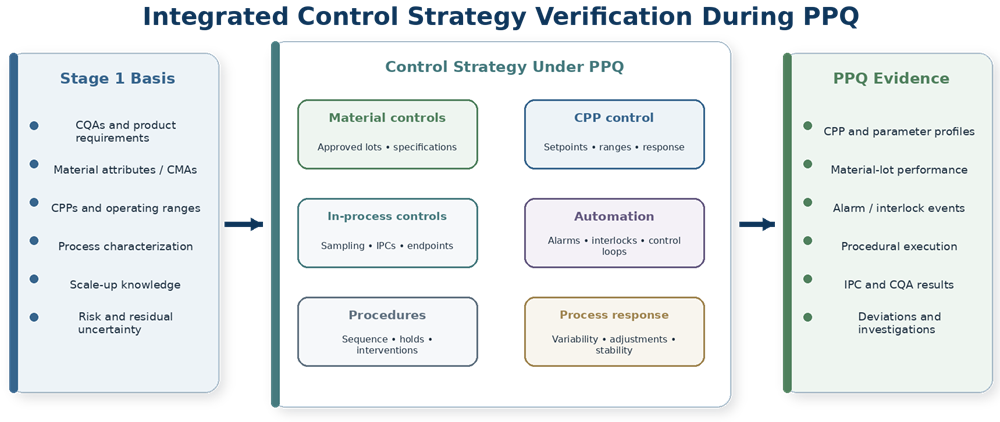
Process Performance Qualification (PPQ) should verify more than whether Critical Process Parameters (CPPs) remained inside predefined limits. The more important Stage 2 question is whether the integrated process control strategy developed during Stage 1 actually works under commercial manufacturing conditions. That requires evaluation of material variability, process parameters, operating ranges, in-process controls, automated and procedural controls, process responses, Critical Quality Attributes (CQAs), deviations, and the consistency of performance across PPQ batches.
FDA’s Process Validation: General Principles and Practices describes PPQ as the combination of qualified facilities, utilities, and equipment with trained personnel, commercial procedures, components, and the commercial manufacturing process. FDA further states that the PPQ approach should reflect overall product and process understanding and demonstrable control, with cumulative development and scale-up evidence used to establish the commercial manufacturing conditions evaluated during PPQ.
The objective is therefore not to repeat Stage 1 development studies at commercial scale. PPQ should confirm that the relationships and controls established during development remain valid when the actual commercial process is operated using routine equipment, materials, personnel, procedures, and supporting systems.
For the Stage 1 basis of these controls, see Process Control Strategy and Design Space Development and CQA, CPP, and Material Attribute Risk Assessment.
PPQ Verifies an Integrated Control Strategy
A process control strategy is a coordinated set of controls derived from product and process understanding. ICH Q8(R2) describes control strategy in terms that can include material attributes, process parameters, equipment operating conditions, in-process controls, finished-product specifications, and the associated monitoring methods and frequencies. It also defines a CPP as a process parameter whose variability can affect a CQA and therefore should be monitored or controlled.
This means that verification during PPQ should not be organized around CPPs alone. A commercial process may remain within every formal CPP limit while still demonstrating poor control through excessive adjustments, unexpected material sensitivity, recurring alarms, unstable process responses, procedural dependence, or unfavorable CQA trends. Conversely, normal variation in a CPP within its approved range is not a problem merely because the parameter is classified as critical; the control strategy should allow expected variation while maintaining acceptable process and product performance.
The PPQ evaluation should therefore answer three connected questions: Were the defined controls implemented as intended? Did they maintain predictable process performance? Did the resulting process consistently produce acceptable quality?
Commercial Manufacturing Conditions Matter
FDA expects the commercial manufacturing process and routine procedures to be followed during PPQ. PPQ lots should be manufactured under normal operating conditions using the personnel routinely expected to perform the process, with representative utilities, materials, environment, equipment, and manufacturing procedures.
This requirement is important because control-strategy verification is meaningful only if the strategy being evaluated is essentially the one intended for routine production. Extraordinary engineering support, atypical material selection, temporary process controls, unusual operator supervision, or repeated manual correction can make an apparently successful PPQ campaign poorly representative of future manufacturing.
Enhanced sampling, additional observation, and increased data collection are appropriate during PPQ. Artificially making the manufacturing process easier to control than it will be during routine production is not.
Stage 1 Evidence Defines What PPQ Must Verify
PPQ should begin with clearly documented Stage 1 conclusions. These normally include identified CQAs, relevant material attributes, CPPs and other important parameters, proposed operating ranges, process-control mechanisms, process interactions, scale-up considerations, hold times, in-process controls, specifications, and remaining uncertainty.
The PPQ protocol should translate those conclusions into specific commercial-scale verification objectives. A parameter identified as sensitive during Design of Experiments (DOE) may require detailed profile evaluation; a material attribute known to affect process behavior may require lot-to-lot comparison; an automated control loop may require confirmation of stable operation; a procedural control may require evidence that trained operators can execute it consistently under routine conditions.
See Process Performance Qualification (PPQ) Strategy and Batch Selection for the overall PPQ strategy and PPQ Sampling Plan and Data Collection Strategy for development of the evidence-collection plan.
Verification of CPPs
CPP verification begins with confirming that parameter values remain within their approved operating ranges, but it should not end there. The evaluation should examine the profile and behavior of each relevant CPP within individual batches and across the PPQ campaign.
Depending on the process, the assessment should consider parameter centering, variability, drift, start-up behavior, transitions, end-of-process behavior, adjustments, excursions, control-loop activity, equipment differences, and reproducibility among batches. A temperature parameter that remains within a 50–60°C range, for example, may still deserve investigation if one batch repeatedly operates around 59.8°C while others remain near the intended setpoint.
The evaluation should distinguish a parameter excursion from normal parameter variation. A well-controlled process is not expected to produce perfectly flat parameter profiles; the relevant question is whether observed variation is predictable, appropriately controlled, and consistent with the process understanding established during development.
Operating Ranges Should Be Evaluated in Context
PPQ should confirm that the routine commercial operating ranges established during Stage 1 are suitable at scale. This does not mean deliberately forcing every CPP to its upper and lower development limits during commercial PPQ.
FDA specifically notes that it is not typically necessary to explore the entire operating range at commercial scale when adequate assurance has already been generated through process-design data.
Commercial PPQ should instead determine whether the process operates appropriately within the intended control strategy and whether naturally occurring or deliberately justified routine variation remains consistent with Stage 1 conclusions. Where PPQ includes operation near a particular range boundary, the rationale should be connected to process risk, commercial representativeness, or remaining uncertainty rather than a generic requirement to “challenge all limits.”
Material Attributes Are Part of the Control Strategy
Material attributes should be evaluated alongside process parameters because incoming material variability can influence process performance even when all CPPs remain controlled. Relevant characteristics may include particle size, moisture, viscosity, concentration, purity, potency, morphology, density, bioburden, supplier-related characteristics, or other attributes identified during development.
PPQ should use representative commercial materials and approved suppliers. Where the PPQ strategy intentionally includes different material lots or suppliers, the evaluation should determine whether observed material variation alters process response, parameter behavior, in-process results, or CQAs.
It is usually unnecessary to force every Critical Material Attribute (CMA) to an artificial extreme during PPQ. Stage 1 characterization should already have established the material–process relationship; PPQ confirms that the material-control strategy remains effective with representative commercial inputs.
CPP–CQA Relationships During PPQ
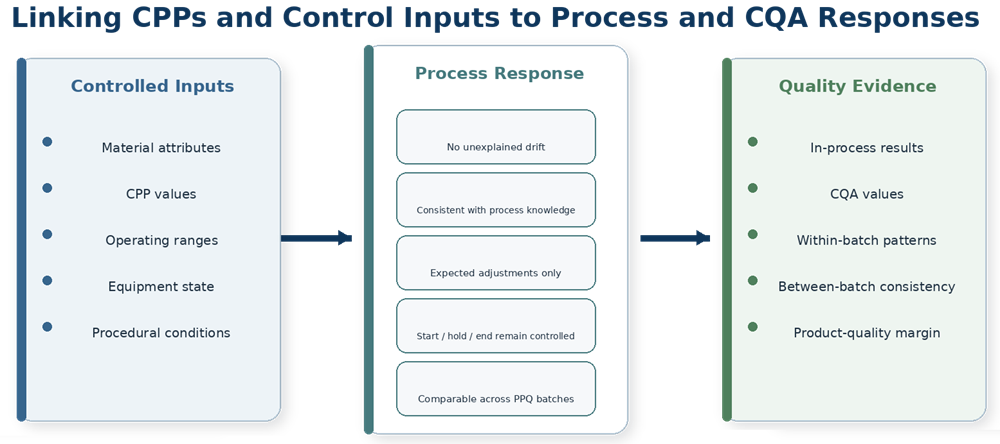
The purpose of evaluating CPP and CQA data together is not to recreate the complete causal model developed during Stage 1. PPQ provides commercial-scale confirmation that the process behaves consistently with that established model.
For a CPP, the analysis should determine whether its observed commercial variation remains compatible with acceptable process responses and CQAs. If different portions of the approved CPP range systematically correspond to shifts in an important CQA, the result may indicate less operating margin than expected even when every individual result remains within specification.
Likewise, a CQA result should not be interpreted only as pass or fail. Patterns within specification can be informative. Movement toward a specification limit, increasing variability, batch-to-batch shifts, or relationships with particular materials or process conditions can challenge assumptions about process robustness.
Process Responses Provide Important Evidence
Many processes generate meaningful responses before final CQAs are measured. These may include endpoint measurements, yield, pressure differential, mixing response, filtration flux, drying rate, process time, equipment load, torque, moisture trajectory, pH adjustment behavior, fill-weight stability, or other indicators of process performance.
These responses can reveal deterioration or control weakness that would be missed by reviewing only final-product results. If a process repeatedly requires longer processing, larger adjustments, more interventions, or greater control effort to achieve the same finished-product result, the control strategy may not be performing as robustly as intended.
PPQ should therefore consider both quality outcomes and how the process achieved those outcomes.
In-Process Controls
21 CFR 211.110 requires written procedures describing in-process controls and appropriate tests or examinations used to monitor output and validate the performance of manufacturing processes that may cause variability.
In-process controls (IPCs) may include physical measurements, analytical tests, visual observations, automated measurements, or intermediate acceptance criteria. During PPQ, the organization should evaluate whether these controls occur at the intended process stage, use representative samples, provide reliable information, and support the process decisions for which they were designed.
An IPC that is consistently acceptable but provides no useful discrimination may warrant later reconsideration. Conversely, an IPC that repeatedly requires adjustment or frequently approaches its acceptance boundary may identify a control-strategy weakness even when the final product remains acceptable.
Alarms and Interlocks
Alarms and interlocks should be included in PPQ evaluation where they are part of the commercial process control strategy. Their full functional challenge normally belongs to equipment or computerized-system qualification rather than being unnecessarily repeated during every PPQ campaign.
PPQ should instead confirm that the required alarm and interlock functions are available in the released configuration and evaluate any alarms, interlocks, overrides, bypasses, trips, or operator responses that occur during commercial process execution. Recurrent nuisance alarms, frequent alarm acknowledgements, repeated excursions that do not trigger expected alarms, or routine dependence on bypass conditions can indicate that the implemented control strategy differs materially from the design intent.
21 CFR 211.68 requires automatic, mechanical, and electronic equipment used in drug manufacturing to be routinely calibrated, inspected, or checked according to written programs designed to assure proper performance, while computer-related systems require appropriate controls over regulated records and functions.
Control Loops and Automated Adjustments
A CPP may remain inside its limits because an automated control loop continuously adjusts the process. That can represent effective process control, but PPQ should determine whether the control action itself is reasonable and consistent with Stage 1 expectations.
Useful evidence may include controller output, frequency and magnitude of adjustments, oscillation, response time, valve or actuator behavior, sensor stability, and interaction with other process variables. Excessive control activity can indicate poor tuning, inadequate equipment capability, unexpected disturbances, or a narrow practical operating margin.
The important point is that the final controlled value alone does not describe the complete process behavior.
Procedural Controls and Operator Execution
Not every control is automated. Stage 1 control strategies may depend on manual additions, sampling, equipment setup, order of operations, hold-time management, transfer sequence, visual checks, defined interventions, or operator responses to process conditions.
PPQ should demonstrate that these procedural controls can be performed consistently by trained routine personnel. FDA requires PPQ lots to be manufactured using routine procedures and normal personnel, and 21 CFR 211.100 requires production and process-control procedures to be followed in execution; deviations from those procedures must be recorded and justified.
Heavy dependence on individual operator technique, frequent clarification of instructions, repeated timing errors, or recurring procedural departures may indicate that the process design or procedures are insufficiently robust. The conclusion should not be masked merely because subsequent testing produced acceptable product.
Hold Times and Process Transitions
Hold times and transitions can be important parts of the control strategy because the material may continue changing when active processing stops. PPQ should confirm that intended commercial holds, transfers, delays, and process transitions remain within the justified strategy and do not introduce unexpected variability.
Relevant evidence can include actual hold duration, temperature, agitation status, storage condition, product or intermediate response before and after the hold, and downstream process behavior. If actual PPQ holds are consistently much shorter than the normal commercial maximum, the campaign may provide limited confirmation of a hold-time control that will be important during routine production.
The appropriate extent of coverage depends on the Stage 1 evidence and PPQ strategy rather than an expectation that every PPQ batch must reach the maximum validated hold.
Equipment and Scale Effects
The control strategy should function on the actual commercial equipment. PPQ therefore provides an important confirmation of scale-dependent assumptions concerning mixing, heat transfer, mass transfer, filtration behavior, residence time, equipment loading, control-loop response, sensor location, process duration, and other equipment–process interactions.
Equipment qualification demonstrates that the equipment can perform its required functions. PPQ determines whether that qualified capability is sufficient when integrated into the actual product-specific commercial process.
Unexpectedly slow heating, increased mixing time, different control-loop behavior, changed filtration profile, greater within-batch variation, or frequent operator intervention may indicate that scale-up knowledge was incomplete even when individual equipment qualification tests were successful.
Do Not Reperform OQ During PPQ
Operational Qualification (OQ) and PPQ serve different purposes. OQ challenges equipment functions, operating ranges, alarms, interlocks, sequences, sensors, and failure responses under controlled qualification conditions. PPQ evaluates the actual commercial manufacturing process using qualified systems and routine procedures.
Repeating every OQ alarm or equipment challenge during PPQ adds little process evidence and can make the PPQ campaign less representative. The PPQ protocol should instead identify which control functions need commercial-process confirmation and which are adequately supported by prior equipment qualification.
This distinction keeps Stage 2 process validation focused on integrated process performance rather than duplicating equipment qualification.
Within-Batch Evaluation
Control-strategy effectiveness should be evaluated across the complete batch where process behavior can change with time, location, equipment condition, material depletion, or process sequence. CPP profiles, IPCs, process responses, and CQA samples should therefore be aligned with the sampling framework established in PPQ Sampling Plan and Data Collection Strategy.
Beginning, middle, and end comparisons can be useful for operations in which process progression is relevant, but the evaluation should remain process-specific. Other processes may require top/middle/bottom comparisons, equipment-zone analysis, pre/post-hold assessment, filling-head comparison, or another scientifically meaningful stratification.
The objective is to determine whether the control strategy performs consistently throughout the batch rather than relying on a batch average that could conceal localized variability.
Between-Batch Evaluation
Each PPQ batch should also be compared with the other qualification batches. Consistent final specifications are necessary, but the comparison should include process-parameter profiles, material characteristics, in-process results, process responses, alarms, adjustments, yields, CQA distributions, and relevant deviations.
Batch-to-batch differences should be interpreted rather than automatically treated as failures. Normal variation is expected; the important question is whether differences are explainable and remain consistent with process understanding.
A campaign in which each batch follows a materially different process trajectory but eventually reaches acceptable finished-product results provides weaker evidence of reproducibility than a campaign with predictable, comparable process behavior.
Control Strategy Verification Matrix
A compact verification matrix can make the PPQ protocol and report more traceable:
| Control element | PPQ question | Typical evidence |
|---|---|---|
| Material attributes | Are representative materials adequately controlled? | Lot data, supplier, material attributes, process response |
| CPPs | Are parameters controlled and reproducible? | Profiles, ranges, trends, adjustments |
| Operating ranges | Are routine ranges suitable commercially? | Actual operating distribution, boundary proximity |
| In-process controls | Do IPCs provide reliable and useful control? | Sample results, timing, response to results |
| Automation | Do control loops perform as intended? | Sensor trends, controller output, adjustments |
| Alarms/interlocks | Do defined safeguards support routine control? | Events, acknowledgements, trips, response |
| Procedural controls | Can routine personnel execute consistently? | Batch records, timing, interventions |
| Process response | Is behavior stable and predictable? | Yield, endpoints, process trajectories |
| CQAs | Is product quality consistently achieved? | Individual results, distributions, trends |
| Deviations | Do unexpected events challenge the strategy? | Investigations, impact assessment, CAPA |
The matrix should trace back to Stage 1 process knowledge rather than create new classifications during PPQ.
Deviations Are Control-Strategy Evidence
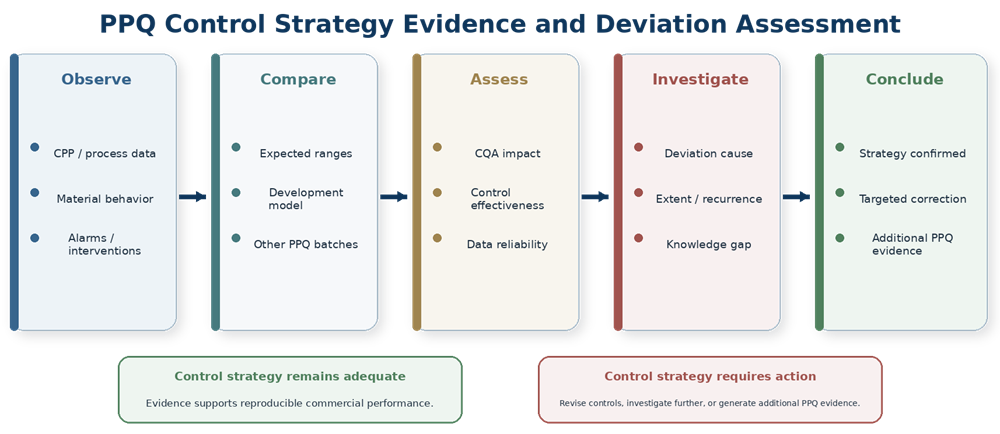
A PPQ deviation should be evaluated for what it reveals about the process. FDA expects the PPQ protocol to include provisions for deviations and nonconforming data and specifically states that PPQ data should not be excluded without documented, science-based justification.
For example, a deviation may demonstrate that a procedure is difficult to execute, an alarm setpoint is poorly positioned, material variability was underestimated, a process range lacks sufficient margin, an equipment response differs at scale, or an assumed CPP–CQA relationship is incomplete. Simply correcting the immediate batch record without considering the control-strategy implication can miss the most important validation information.
The investigation should determine whether the event was isolated, whether it affected product quality or process interpretability, whether it can recur under routine manufacture, and whether the existing strategy remains adequate.
Frequent Adjustments Can Indicate Weak Control
A common analytical mistake is to conclude that a CPP is well controlled simply because every recorded value remained inside specification or operating limits. If operators or control systems required repeated correction to prevent excursions, the apparent compliance may reflect active recovery rather than inherent process stability.
The PPQ report should therefore consider the number and magnitude of manual adjustments, automated corrections, interventions, alarm responses, and atypical operator actions. Frequent correction does not automatically mean the process has failed, but it should be explained and compared with the intended commercial control philosophy.
A robust process should generally demonstrate predictable operation with a level of intervention consistent with the process design.
Acceptance Criteria Should Reflect Integrated Performance
CPP verification criteria should not be reduced to “all CPPs within limits.” Similarly, control-strategy verification should not be reduced to “all CQAs met specification.”
The PPQ conclusion should integrate process and product evidence. Depending on the process, acceptable performance may require that CPPs remain appropriately controlled, material variability is accommodated, IPCs function as intended, process responses are predictable, alarms and interventions are not excessive, procedural execution is consistent, CQAs meet requirements with appropriate margin, and no unresolved event contradicts the expected state of control.
Detailed statistical and acceptance methodology belongs in PPQ Acceptance Criteria and Statistical Evaluation.
Statistical Evaluation
Statistical evaluation can help determine whether observed parameter and response variation is consistent within and between PPQ batches. FDA specifically recommends that the PPQ protocol define statistical methods capable of evaluating intra-batch and inter-batch variability.
Depending on the process and data type, useful methods can include descriptive statistics, control or run charts, comparison of distributions, variance analysis, regression, parameter–response analysis, confidence intervals, or capability/performance measures where justified. Statistics should support scientific interpretation; an impressive model cannot compensate for an unrepresentative PPQ campaign or incomplete process knowledge.
The most useful question is often not whether a statistical test returned a particular p-value, but whether the data demonstrate stable and reproducible process behavior consistent with the control strategy.
When PPQ Challenges Stage 1 Conclusions
PPQ can reveal that a Stage 1 assumption was incomplete. This is not necessarily a validation failure; discovering and addressing the issue is part of the lifecycle model.
Examples include a parameter behaving more sensitively at commercial scale, an approved material attribute causing greater process variation than expected, an operating range proving too broad, control-loop performance differing from development equipment, or a procedural control proving difficult to execute consistently. The response should be proportionate to the significance of the new evidence.
Possible actions include revising a parameter range, strengthening a material control, modifying an alarm, changing a procedure, performing targeted characterization, updating the risk assessment, or generating additional PPQ evidence. Significant changes should be formally controlled and their effect on existing PPQ batches assessed.
PPQ Report Conclusion
The PPQ report should state whether the integrated control strategy has been confirmed under commercial manufacturing conditions. The conclusion should explain how Stage 1 assumptions were evaluated and whether the resulting process data, material information, IPCs, automated and procedural controls, CQAs, deviations, and statistical analysis collectively support reproducible process performance.
A useful final conclusion should answer whether the strategy is adequate as designed, whether minor refinements are required without challenging the overall PPQ conclusion, or whether the evidence requires more substantial process-development or qualification activity. It should also identify which controls and process relationships require continued monitoring during Stage 3.
FDA expects the PPQ report to summarize and analyze collected data, address protocol adherence and deviations, discuss corrective actions or process changes, and determine whether the process is in a state of control.
Transition to Continued Process Verification
Successful PPQ does not freeze CPP classifications, parameter ranges, alarms, monitoring strategies, or procedural controls permanently. Stage 3 Continued Process Verification (CPV) provides a much larger commercial dataset and may confirm or challenge the original control-strategy conclusions.
The PPQ report should therefore identify the parameters, material attributes, process responses, IPCs, CQAs, or control-system behaviors that warrant continued monitoring. Particular attention may be appropriate where PPQ identified limited operating margin, material sensitivity, unusual variability, frequent adjustments, residual uncertainty, or follow-up actions.
See Continued Process Verification (CPV) Program and Monitoring Strategy for the Stage 3 monitoring framework.
Key Principles
- PPQ verifies the integrated process control strategy, not CPP limits in isolation.
- CPPs should be evaluated for profile, variability, drift, reproducibility, and relationship to process and quality responses.
- PPQ does not normally need to challenge the complete development range at commercial scale when Stage 1 evidence already provides adequate assurance.
- Material attributes and representative material variability are part of control-strategy verification.
- CPP–CQA relationships should be confirmed as commercially consistent with Stage 1 understanding rather than redeveloped from scratch.
- In-process controls should provide meaningful evidence about process state or intermediate quality.
- Full functional alarm and interlock challenge belongs principally to qualification; PPQ evaluates their role in the actual commercial process.
- Frequent manual or automated corrections may indicate limited process robustness even when all values remain within limits.
- Procedural controls should be demonstrated using routine trained personnel and approved production procedures.
- Within-batch and between-batch evidence should be evaluated together.
- Deviations should be treated as information about process control and should not be discarded merely because the batch ultimately meets specification.
- Final PPQ acceptance should integrate material, process, control-system, in-process, CQA, statistical, and deviation evidence.
- Stage 3 monitoring should continue to confirm that the control strategy remains effective.