

PPQ Strategy and Batch Selection
Process Performance Qualification (PPQ) is the commercial-scale demonstration that the manufacturing process defined during Stage 1 can perform reproducibly under the conditions intended for routine production. PPQ is not simply the manufacture of a predetermined number of successful batches. It is a structured evaluation of the integrated commercial process, its variability, and the effectiveness of its control strategy.
FDA’s Process Validation: General Principles and Practices places PPQ within Stage 2 — Process Qualification. FDA distinguishes two elements of Stage 2: qualification of the facility, utilities, and equipment, followed by PPQ of the actual commercial manufacturing process. PPQ combines the qualified facility and systems with trained personnel, commercial procedures, materials, controls, and the intended manufacturing process to produce commercial-scale batches.
A successful PPQ provides evidence that the process design developed during Stage 1 performs as expected at commercial scale. The strategy should therefore be based on accumulated process knowledge, risk, known sources of variability, commercial-scale considerations, and the strength of the available evidence—not an inherited assumption that three successful batches automatically establish validation.
For the overall lifecycle framework, see General Principles of Process Validation and Regulatory Requirements and FDA Expectations for Process Validation.
PPQ Within Stage 2 Process Qualification
PPQ — Process Performance Qualification is the second element of FDA Stage 2 Process Qualification. It should not be confused with equipment Performance Qualification.
Equipment qualification demonstrates that facilities, utilities, and equipment are suitable for their intended use and can operate within required ranges and loads. PPQ evaluates the performance of the integrated manufacturing process using those qualified systems.
FDA states that facility design, commissioning, and qualification of utilities and equipment should precede PPQ. The equipment should be capable of performing under loads comparable to routine production and under anticipated operating conditions, including expected interventions, stoppages, start-ups, and operating durations.
The progression is therefore: Stage 1 Process Design → Facility/Utility/Equipment Qualification → PPQ → Stage 3 Continued Process Verification
PPQ should not be initiated merely because equipment IQ/OQ/PQ documentation is complete. The manufacturing process itself must also be sufficiently mature and defined.
PPQ Readiness
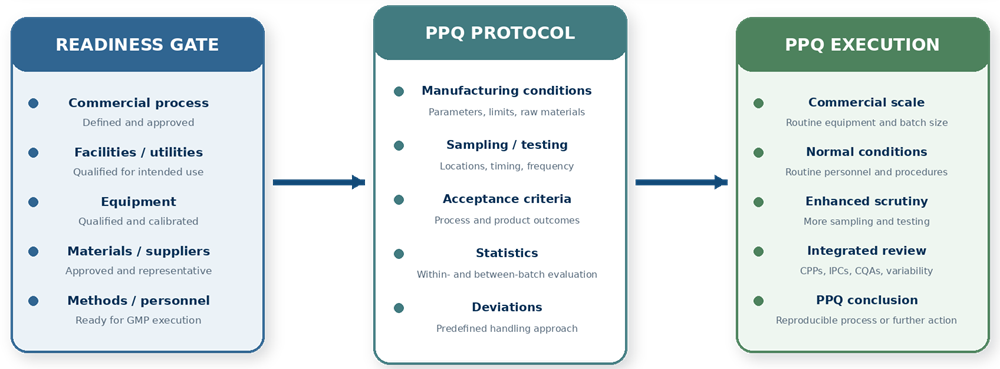
Before PPQ begins, the organization should establish that the process and manufacturing environment are ready for meaningful commercial-scale evaluation. Typical readiness prerequisites include:
- the commercial process has been defined;
- Stage 1 characterization is substantially complete;
- Critical Quality Attributes (CQAs) have been established;
- Critical Process Parameters (CPPs) and other important parameters have been evaluated;
- the process control strategy has been defined;
- commercial operating ranges and setpoints are established;
- commercial batch size or batch-size range is defined;
- relevant scale-up relationships have been evaluated;
- facilities and utilities are qualified;
- manufacturing equipment is qualified for intended use;
- calibration and maintenance status is acceptable;
- raw materials, components, and suppliers are approved;
- material attributes relevant to process performance are understood;
- manufacturing and cleaning procedures are approved;
- operators are trained and qualified;
- analytical methods are suitable for PPQ and commercial testing;
- sampling and testing capability is available;
- computerized or automated systems supporting the process are ready for GMP use;
- deviations and investigations can be managed through established procedures; and
- the PPQ protocol has been reviewed and approved.
FDA specifically identifies facility and equipment qualification, personnel training, material-source verification, analytical-method status, and appropriate departmental and Quality Unit approval among PPQ protocol considerations. PPQ should not become the mechanism for closing major unresolved Stage 1 development gaps.
The PPQ Strategy
The PPQ strategy defines how sufficient evidence will be generated to determine whether the commercial process performs reproducibly. It should be developed from Stage 1 knowledge rather than beginning with the question, “How many batches do we need?”
The more appropriate sequence is: What must be demonstrated? → What variability must be evaluated? → What data are required? → Which batches and conditions provide that evidence?
Important strategy inputs include:
- development and characterization results;
- Design of Experiments (DOE);
- process risk assessments;
- CQA–CPP relationships;
- material variability;
- scale-up knowledge;
- equipment/process interactions;
- commercial operating ranges;
- process robustness;
- hold-time studies;
- historical manufacturing experience;
- prior experience with sufficiently similar processes;
- residual uncertainty; and
- the established control strategy.
FDA explicitly recommends using cumulative information from designed experiments, laboratory studies, pilot work, commercial experience, and other relevant studies to establish PPQ manufacturing conditions.
See Process Characterization and Development Studies for Process Validation, Design of Experiments for Process Characterization and Validation, and Process Control Strategy and Design Space Development for the Stage 1 evidence that should feed into PPQ.
The PPQ Protocol
FDA describes a written PPQ protocol defining manufacturing conditions, controls, testing, and expected outcomes as essential to Stage 2. The protocol should define, as applicable:
- product and process scope;
- commercial manufacturing site;
- manufacturing equipment and utilities;
- batch size;
- number and sequence of PPQ batches;
- material and component requirements;
- relevant material attributes;
- process parameters and operating ranges;
- CPPs;
- in-process controls;
- manufacturing conditions;
- process limits;
- sampling locations;
- sampling timing;
- number of samples;
- sampling frequency;
- release testing;
- additional PPQ characterization testing;
- CQA evaluation;
- statistical methods;
- within-batch analysis;
- between-batch analysis;
- predefined acceptance criteria;
- process-performance indicators;
- deviation and nonconformance handling;
- protocol departure requirements;
- overall PPQ success criteria;
- responsibilities; and
- review and approval requirements.
The protocol should describe how the complete dataset will support the final process-performance conclusion rather than establishing disconnected pass/fail criteria for individual observations.
See PPQ Acceptance Criteria and Statistical Evaluation for the detailed evaluation framework.
PPQ Must Use the Commercial Process
PPQ should represent the process intended for routine manufacturing. FDA expects PPQ lots to be manufactured using the commercial manufacturing process and routine procedures under normal conditions by the personnel expected to perform routine manufacturing. Normal conditions include utilities, materials, personnel, environment, and manufacturing procedures.
PPQ batches should therefore use, as applicable:
- commercial-scale equipment;
- commercial batch records;
- routine manufacturing procedures;
- intended material grades and components;
- approved suppliers;
- routine automation;
- intended process setpoints and ranges;
- routine production personnel;
- normal utility systems;
- intended hold times;
- normal material transfers; and
- the established control strategy.
A PPQ campaign that requires extraordinary engineering attention, special adjustments, atypical operator intervention, unusually selected materials, or temporary controls not intended for routine manufacturing may provide poor evidence of routine process capability.
This does not mean the PPQ team cannot provide enhanced observation and data collection. Enhanced monitoring is appropriate; artificially improving the manufacturing process for PPQ is not.
Commercial Batch Size
FDA expects the decision supporting commercial distribution to be based on commercial-scale batch data. Laboratory and pilot studies provide supporting knowledge but do not substitute for commercial-scale demonstration.
- Where only one commercial batch size is intended, the PPQ strategy should normally use that defined commercial size.
- Where a commercial batch-size range is intended, the strategy should determine which size or sizes provide sufficient evidence.
This determination should consider process mechanism rather than assuming that the largest batch is automatically worst case. For some operations, maximum batch size can represent increased risk because of:
- mixing limitations;
- heat-transfer limitations;
- mass-transfer limitations;
- longer processing time;
- longer filling duration;
- greater filtration load;
- increased hold time; or
- equipment capacity.
For other operations, the minimum batch size may be more challenging because of:
- minimum equipment loading;
- altered mixing geometry;
- reduced probe immersion;
- different heat-transfer characteristics;
- greater surface-area-to-volume effects; or
- equipment-control limitations.
The PPQ protocol should therefore state why the selected batch size or sizes are representative of the intended commercial process.
PPQ Batch Selection
PPQ batches should be selected prospectively through the approved strategy rather than retrospectively selecting successful manufacturing runs. Batch selection should allow the study to evaluate expected sources of routine variation.
Depending on the process, relevant variation can include:
- raw-material lots;
- component lots;
- approved suppliers;
- equipment trains;
- manufacturing shifts;
- operators;
- normal environmental variation;
- normal process timing;
- expected hold durations;
- equipment loading;
- process sequence;
- intermediate storage; and
- other known sources of commercial variability.
The purpose is not to maximize variation artificially. It is to avoid constructing a PPQ campaign so tightly controlled that it fails to represent routine manufacturing.
FDA Does Not Require Three PPQ Batches
This point should be explicit.
FDA’s CGMP Questions and Answers — Production and Process Controls directly asks whether CGMP requires three successful process-validation batches.
FDA’s answer is no. Neither CGMP regulations nor FDA policy specifies a minimum number of batches required to validate a manufacturing process. FDA also states that process validation cannot be reduced to the simplistic formula of completing three successful full-scale batches and expects the manufacturer to provide a sound rationale for the selected number.
Therefore:
- Three batches may be appropriate.
- Two, four, five, six, or another number may be appropriate.
- The regulatory weakness is not choosing three—it is choosing three without a process-specific scientific rationale.
The same applies to the concept of “three consecutive successful batches.” FDA does not establish universal consecutiveness as a regulatory requirement.
Establishing the Number of PPQ Batches
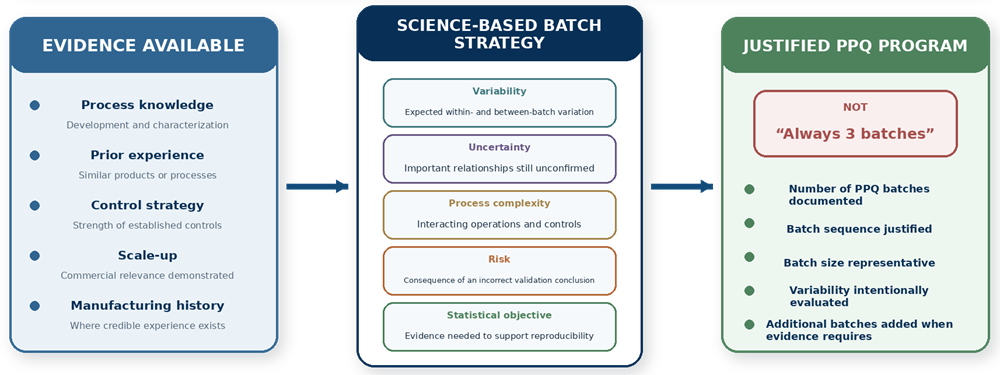
The number of PPQ batches should be sufficient to support a high degree of assurance that the commercial process is reproducible. Important considerations include:
Process Knowledge
A highly characterized process with strong mechanistic understanding and extensive development evidence may require a different PPQ strategy than a process with substantial unresolved uncertainty.
Process Variability
A process with significant expected batch-to-batch variation generally requires more evidence than a process that has demonstrated highly consistent behavior. Both intra-batch variability—variation within one batch—and inter-batch variability—variation among different batches—should be evaluated.
FDA specifically expects PPQ statistical metrics to address both.
Process Complexity
Complex processes containing multiple interacting unit operations, sensitive biological transformations, significant material dependence, manual interventions, or complex control loops may warrant broader PPQ evidence.
Product and Process Risk
The consequence of incorrectly concluding that an inadequately understood process is validated should influence the strength of evidence required.
Stage 1 Evidence
Strong DOE, characterization, scale-up, and process-history data can reduce uncertainty entering PPQ. Weak or poorly representative development evidence increases it.
Control-Strategy Maturity
A well-understood automated control strategy with reliable measurements may provide stronger assurance than a process dependent on manual control and end-product testing.
Previous Manufacturing Experience
FDA recognizes credible prior experience with sufficiently similar products and processes as potentially useful in determining PPQ strategy. Similarity should be scientifically demonstrated rather than assumed.
Statistical Objectives
The strategy should consider how much data are necessary to evaluate:
- expected within-batch variation;
- between-batch variation;
- process centering;
- reproducibility;
- parameter–CQA relationships;
- confidence in observed performance; and
- residual uncertainty.
No single universal statistical formula converts these inputs into a required PPQ batch count.
A Practical Batch-Number Strategy
A useful approach is to establish the batch strategy prospectively in the PPQ protocol. For example, the protocol can define:
- the planned initial number of batches;
- scientific rationale for that number;
- what evidence each batch contributes;
- required coverage of material or operating conditions;
- statistical evaluation across the batches;
- conditions requiring additional PPQ batches; and
- conditions under which the collected evidence is considered sufficient.
This approach is stronger than manufacturing an arbitrary number of batches and deciding afterward whether the results look acceptable.
Additional PPQ evidence may be warranted when:
- observed variability exceeds development expectations;
- significant batch-to-batch differences occur;
- an important material lot behaves differently;
- scale-up assumptions are challenged;
- control performance is inconsistent;
- unexpected parameter–CQA relationships appear;
- deviations affect interpretability;
- statistical uncertainty remains excessive; or
- the original PPQ rationale is no longer supported.
Additional batches should not be viewed as punishment for an imperfect campaign. They are additional evidence required when the initial evidence does not support the intended conclusion.
Consecutive Batches
Manufacturing PPQ batches consecutively can be operationally useful because it reduces uncontrolled changes between qualification runs and demonstrates repeatability over a defined period.
However, consecutive manufacture should not become a substitute for representativeness. For some processes, manufacturing three tightly clustered batches from the same raw-material lot, same shift, same operators, and nearly identical environmental conditions may provide less information about commercial variability than a scientifically planned campaign spanning relevant routine variation.
The protocol should therefore explain whether consecutiveness contributes meaningfully to the validation objective.
Worst-Case Considerations
PPQ should evaluate process conditions that are representative of the intended commercial process, including justified high-risk or challenging conditions where appropriate. Examples might include:
- maximum or minimum intended batch size;
- maximum validated equipment load;
- longest routine hold time;
- high-risk material characteristics within approved limits;
- an approved supplier known to represent greater process variability;
- challenging equipment configuration;
- beginning or end of long manufacturing runs;
- maximum normal process duration; or
- other conditions identified through Stage 1 risk assessment.
However, PPQ should not be converted into a process-development failure study. FDA notes that it is generally not necessary to explore the entire operating range at commercial scale when Stage 1 data provide adequate assurance.
Artificially forcing PPQ to process failure or deliberately operating outside the intended commercial control strategy can reduce rather than improve the relevance of the qualification.
Variability Is the Central PPQ Question
The underlying objective of PPQ is not simply to show that every test result passes. PPQ should demonstrate that the process behaves reproducibly and that observed variability is consistent with process understanding. This means evaluating:
Within-batch variability
Examples include:
- start/middle/end behavior;
- spatial uniformity;
- equipment-location effects;
- filling sequence;
- material depletion;
- process transitions; and
- time-dependent behavior.
Between-batch variability
Examples include:
- batch means;
- variability profiles;
- material-lot effects;
- shift or operator effects;
- equipment differences;
- batch duration; and
- CQA distributions.
FDA’s process-validation guidance repeatedly emphasizes understanding, detecting, and controlling variation throughout the lifecycle. A batch can meet finished-product specifications and still provide evidence of an inadequately controlled process if its variability is inconsistent with the process design.
Enhanced PPQ Sampling
PPQ should normally use more extensive sampling, testing, and process scrutiny than routine commercial production. FDA explicitly states that PPQ will in most cases have a higher level of sampling, additional testing, and greater scrutiny than routine production.
The PPQ sampling plan should define:
- sampling locations;
- sampling times;
- number of samples;
- sample frequency;
- unit operations covered;
- attributes tested;
- relationship to known variability;
- worst-case locations;
- within-batch distribution;
- between-batch comparisons; and
- statistical rationale.
FDA also states that sample numbers should provide sufficient statistical confidence regarding quality both within a batch and between batches.
21 CFR 211.110 requires in-process controls that monitor output and validate performance of manufacturing processes that may cause variability, while 21 CFR 211.160 requires scientifically sound sampling plans and representative samples.
See PPQ Sampling Plan and Data Collection Strategy for detailed sampling design.
Statistical Justification
FDA strongly recommends objective measures, including statistical metrics, where feasible and meaningful during PPQ. The statistical strategy should be defined before execution and should answer the process question rather than merely produce descriptive output. Depending on the process and available data, analysis may include:
- descriptive statistics;
- within-batch comparisons;
- between-batch comparisons;
- distribution assessment;
- trends;
- variance components;
- confidence intervals;
- tolerance intervals;
- process capability or performance metrics where appropriate;
- parameter–CQA relationships; and
- other scientifically justified methods.
The statistical method should be proportionate to:
- data type;
- sample size;
- process complexity;
- expected variability;
- risk;
- study design; and
- the conclusion being supported.
A mathematically sophisticated analysis cannot compensate for an unrepresentative PPQ campaign. Likewise, simply obtaining three acceptable batch averages provides limited statistical evidence regarding process reproducibility.
See PPQ Acceptance Criteria and Statistical Evaluation for detailed treatment.
PPQ Acceptance Should Be Integrated
The final PPQ conclusion should consider the complete evidence. A successful conclusion generally requires evidence that:
- relevant CQAs meet requirements;
- CPPs remain appropriately controlled;
- in-process controls perform as intended;
- the control strategy is effective;
- within-batch variability is acceptable;
- between-batch behavior is reproducible;
- unexpected trends are adequately understood;
- deviations have been appropriately investigated;
- statistical results support the process conclusion; and
- no unresolved information materially undermines confidence in the process.
Meeting finished-product specifications alone is insufficient evidence that PPQ succeeded.
See Verification of CPPs and Process Control Strategy During PPQ for control-strategy confirmation.
PPQ Deviations
Deviation handling should be predefined in the protocol.
FDA states that PPQ data should not be excluded from consideration without documented science-based justification. The final report should discuss manufacturing nonconformances, aberrant results, unexpected observations, corrective actions, and any information relevant to the validity of the process conclusion.
A deviation does not automatically invalidate a PPQ batch.
Its significance depends on:
- root cause;
- effect on product quality;
- effect on process performance;
- representativeness of the batch;
- effect on collected data;
- whether process control was compromised;
- whether the event indicates an unrecognized source of variability; and
- whether the overall qualification conclusion remains supportable.
See PPQ Deviations, Investigation, and Validation Conclusion for the detailed framework.
PPQ Report and Final Conclusion
The PPQ report should evaluate the full campaign rather than merely state that individual batches passed. FDA expects the report to:
- address adherence to the protocol;
- summarize and analyze collected data;
- evaluate unexpected observations;
- discuss deviations and nonconformances;
- describe required corrective actions or process changes;
- determine whether protocol conditions were met;
- determine whether the process is in a state of control; and
- document the scientific basis for approving the process.
The conclusion should be based on accumulated knowledge from Stage 1 through completion of Process Qualification.
Where the evidence does not demonstrate reproducible commercial performance, the correct response may include:
- additional investigation;
- additional characterization;
- control-strategy revision;
- additional PPQ batches;
- targeted equipment or system work; or
- reconsideration of the process design.
The desired outcome is not an approved report. It is a justified conclusion that the commercial process performs reproducibly.
Transition From PPQ to Stage 3
PPQ should establish the transition into Stage 3 — Continued Process Verification (CPV). This transition should define:
- parameters and attributes requiring ongoing monitoring;
- relevant material characteristics;
- initial CPV sampling frequencies;
- statistical monitoring methods;
- known sources of variability;
- residual risks;
- post-PPQ follow-up commitments;
- CAPA effectiveness checks where applicable; and
- conditions requiring escalation.
Importantly, completion of PPQ does not necessarily mean immediately reducing sampling to routine minimum levels.
FDA recommends continuing monitoring and sampling at the level established during Process Qualification until sufficient data exist to generate meaningful variability estimates. Those estimates can then support statistically appropriate and representative routine monitoring frequencies.
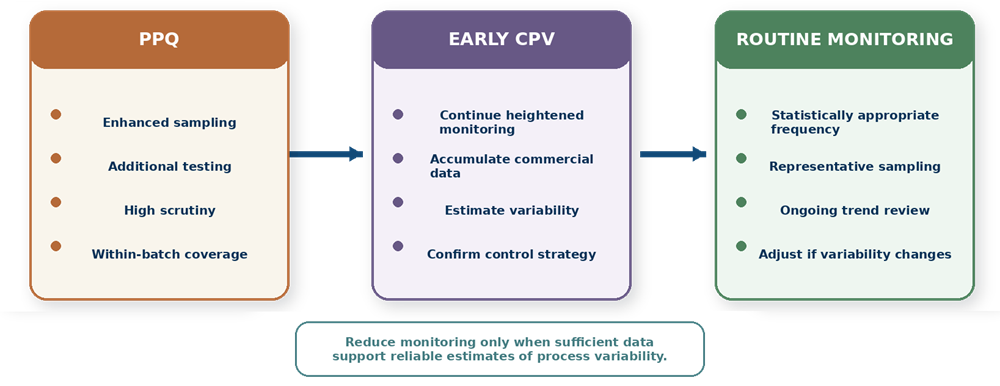
This concept is important because PPQ completion is not a cliff where intensive process knowledge suddenly stops being collected.
See Continued Process Verification (CPV) Program and Monitoring Strategy and Continued Process Verification Reporting and State-of-Control Assessment for Stage 3 lifecycle control.
PPQ and Routine Manufacturing
The transition to routine commercial operation should be deliberate. The organization should have documented evidence that:
- the process design has been confirmed;
- the control strategy performs effectively;
- batch-to-batch reproducibility is demonstrated;
- relevant variability is understood;
- PPQ deviations are resolved or adequately addressed;
- analytical and process data support the qualification conclusion;
- initial CPV requirements are established; and
- responsibilities for ongoing monitoring are clear.
Routine production should then continue generating evidence that the PPQ conclusion remains valid. PPQ is therefore not the end of process validation. It is the point at which accumulated Stage 1 knowledge and commercial-scale Stage 2 evidence support entry into continued lifecycle verification.
Key Principles
- PPQ is part of Stage 2 Process Qualification and is distinct from equipment qualification.
- The commercial process should be defined before PPQ begins.
- PPQ uses qualified facilities, utilities, and equipment, trained personnel, approved procedures, representative materials, and the intended control strategy.
- FDA does not prescribe a universal minimum number of PPQ batches.
- “Three successful batches” is not a universal FDA requirement.
- “Three consecutive batches” is not a universal FDA requirement.
- Batch number should be scientifically justified based on process knowledge, variability, complexity, risk, uncertainty, prior experience, and statistical objectives.
- PPQ batches should represent intended routine commercial manufacturing conditions.
- Commercial batch size or batch-size ranges require scientific justification.
- The largest batch is not automatically worst case.
- Worst-case conditions should remain relevant to the intended commercial process.
- PPQ should not be used as a substitute for incomplete Stage 1 development.
- PPQ normally uses more sampling, testing, and scrutiny than routine production.
- Within-batch and between-batch variability should both be evaluated.
- Statistical methods should be predefined and scientifically appropriate.
- Meeting product specifications alone does not prove adequate process qualification.
- Deviations should be evaluated scientifically rather than automatically invalidating or excluding a PPQ batch.
- Completion of PPQ should establish the transition into CPV.
- Heightened monitoring should continue until sufficient commercial data support statistically appropriate routine monitoring.